
CH 8. Aldehydes, Ketones & Condensations
A. Carbonyl Structure and Reactivity
1. Structure, naming and preparation
a. Carbonyl bonding, geometry and electrophilicity
- C=O contains one σ bond and one π bond. Its carbon is approximately sp²-hybridized and trigonal planar. Polarization toward oxygen leaves carbon partially positive and electrophilic; nucleophilic addition makes it tetrahedral.
- An aldehyde has at least one H on its carbonyl carbon; a ketone has two carbon substituents. Aldehydes are commonly less hindered and more reactive toward addition than comparable ketones.
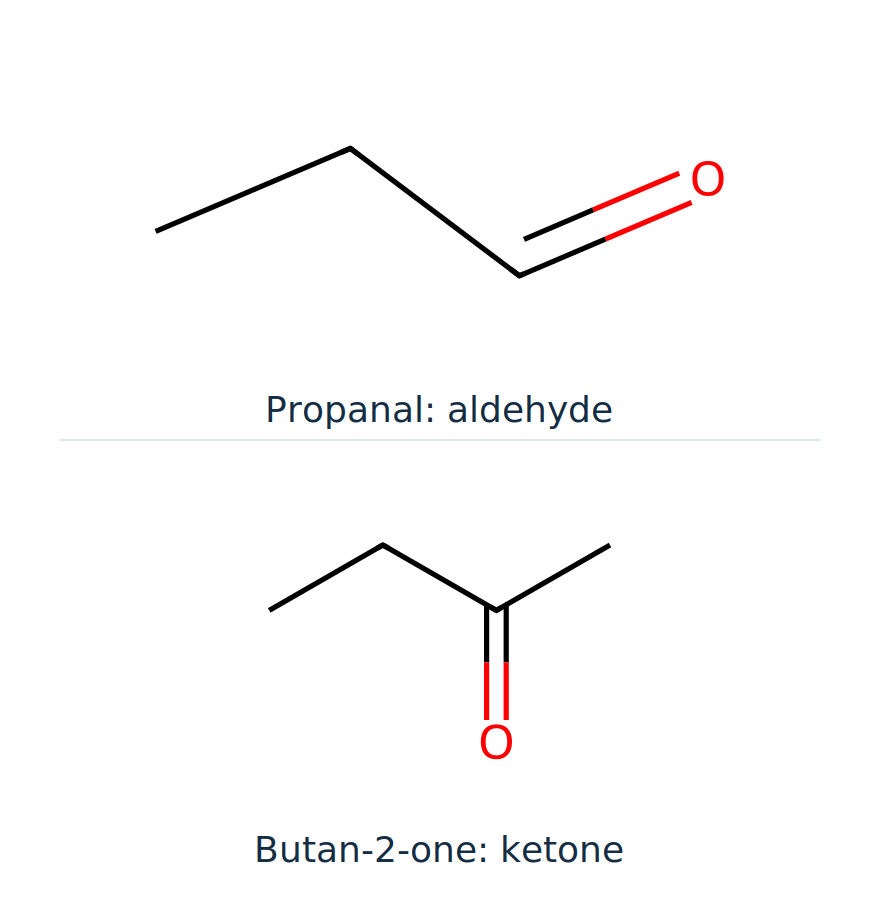
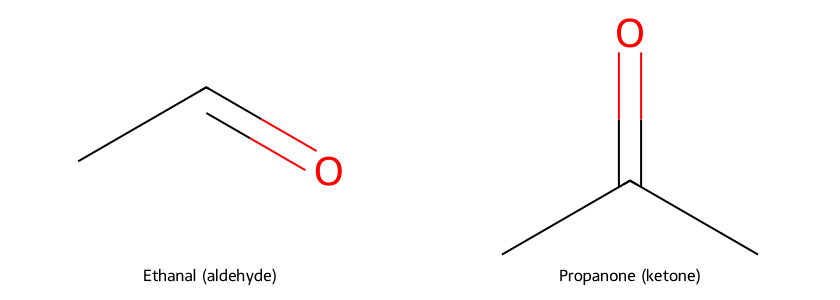
- Lower electron donation as well as lower steric crowding contributes to the greater addition reactivity of many aldehydes.
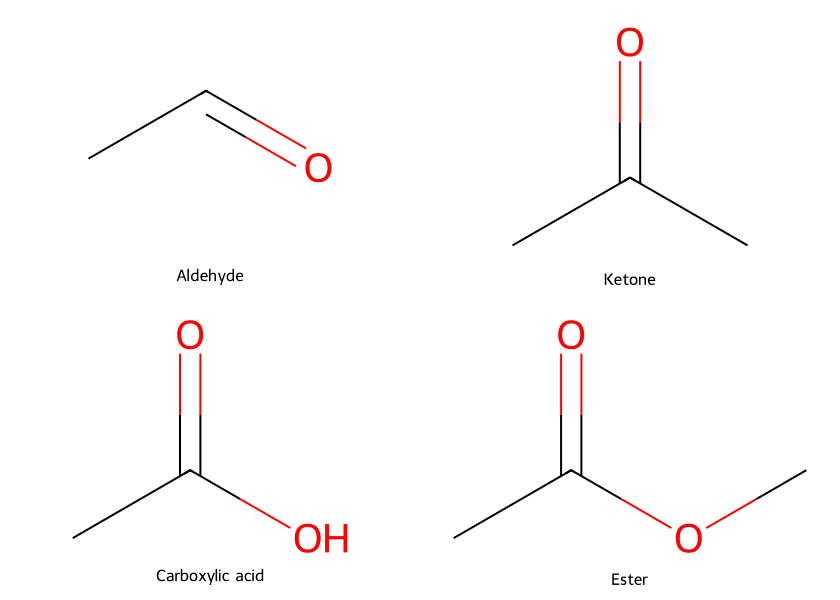
- Related acyl derivatives, such as carboxylic acids and esters, commonly undergo nucleophilic acyl substitution rather than the simple addition characteristic of aldehydes and ketones.
b. Naming aldehydes
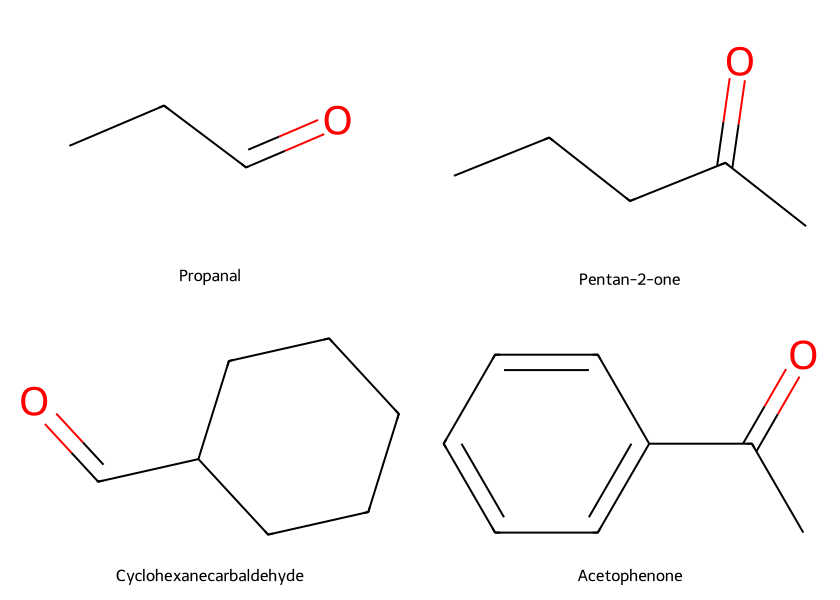
- Replace the terminal -e of the parent alkane with -al for an acyclic aldehyde; the aldehyde carbon is carbon 1.
- When –CHO is directly attached to a ring, the suffix -carbaldehyde is often used.
c. Naming ketones
- Use -one for the ketone suffix and number the longest appropriate chain to give the carbonyl carbon a low locant.
- A carbonyl group written as a substituent is commonly designated by oxo-.
d. Alcohol oxidation
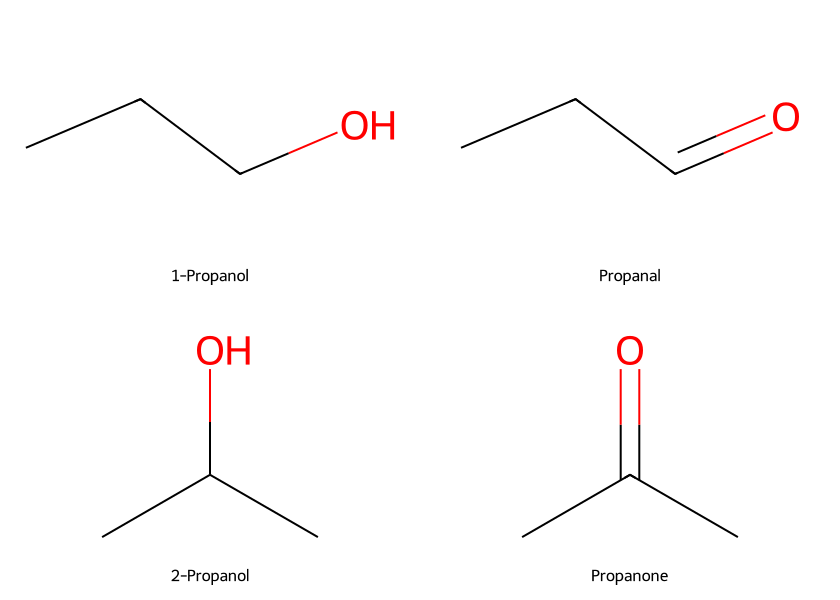
- Primary alcohols can be oxidized to aldehydes with suitable mild conditions, such as PCC; stronger aqueous oxidation can continue to carboxylic acids.
- Secondary alcohols are oxidized to ketones.
- Tertiary alcohols lack an ordinary carbinol hydrogen and do not undergo comparable simple oxidation without C–C bond cleavage.
e. Alternative sources of carbonyl groups
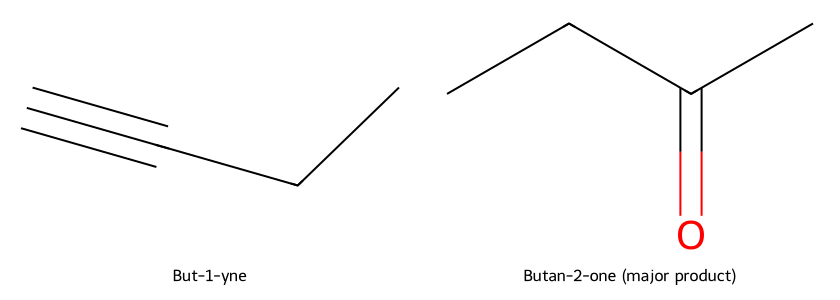
- Acid-catalyzed hydration of a terminal alkyne generally gives a methyl ketone after enol–ketone tautomerization; ethyne gives ethanal.
- Friedel–Crafts acylation can form aryl ketones from aromatic substrates under suitable conditions.
- Alkene cleavage is another route to carbonyl compounds. A formyl group can be named as a substituent when a higher-priority functional group controls the suffix.
B. Nucleophilic Addition
1. Acidic and basic addition pathways
a. Basic or nucleophilic conditions
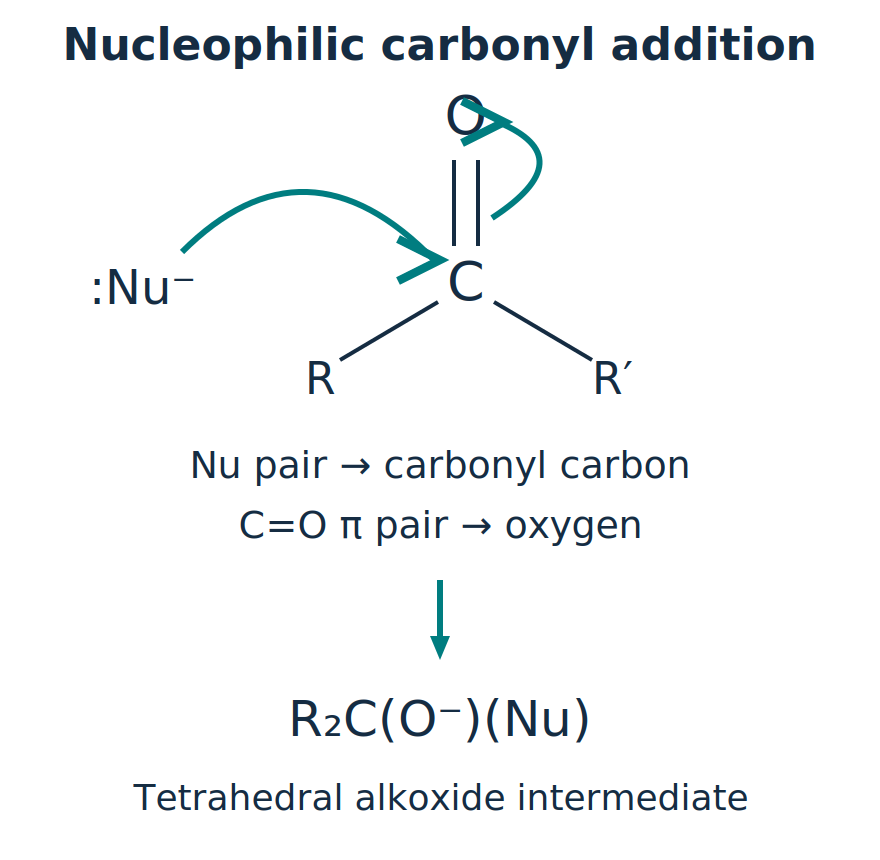
- A nucleophile supplies an electron pair to the electrophilic carbonyl carbon.
- The carbonyl π electrons move toward oxygen as a tetrahedral alkoxide intermediate forms.
- Protonation converts the alkoxide to an alcohol.
- Aldehydes are often more reactive than structurally comparable ketones because steric crowding and electron donation are generally lower.

- Hydride, organometallic carbon and cyanide provide different nucleophiles; in each case the new bond forms to carbonyl carbon rather than oxygen.
b. Acid-catalyzed addition
- Protonation of carbonyl oxygen increases electrophilicity at the carbonyl carbon.
- A neutral nucleophile, such as water or an alcohol, may then attack; deprotonation gives the addition product.
- The acid-catalyzed and basic pathways are distinct and must not be drawn using the wrong protonation states.
- Strongly basic organometallic reagents are incompatible with aqueous acid during addition; introduce acid only in a separate workup.
c. Carbonyl hydrate
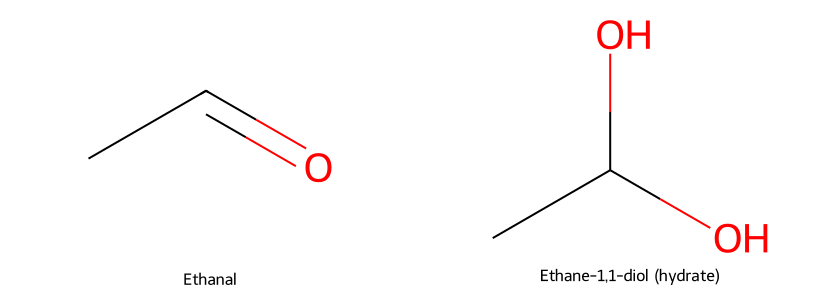
- Water adds reversibly across the C=O group to give a geminal diol (two –OH groups on the same carbon).
- Hydration can be accelerated by acid or base; the equilibrium position depends strongly on substrate substitution and electron-withdrawing effects.
- Formaldehyde and some strongly electron-poor aldehydes have relatively favorable hydration equilibria compared with typical ketones.
d. Cyanohydrin formation
- Cyanide can add to a carbonyl group, and protonation gives a cyanohydrin containing –OH and –CN on the same carbon.
- Cyanohydrin formation creates a new C–C bond; the new alcohol-bearing carbon may become stereogenic.
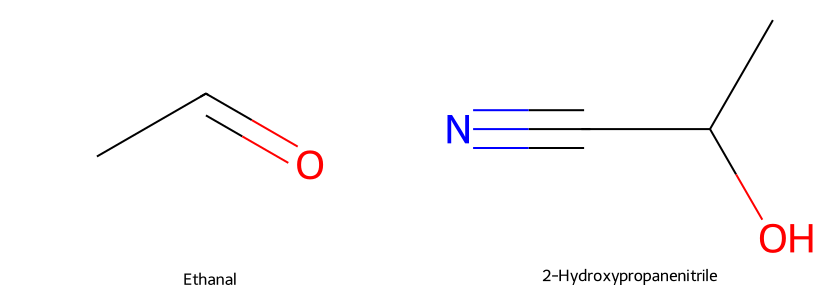
- Formation is reversible and depends on substrate and conditions. Later nitrile hydrolysis can give a carboxylic acid; include the carbon introduced by CN in the product carbon count.

C. Grignard and Hydride Addition
1. Carbon–carbon bonds and reduction products
a. NaBH₄ and LiAlH₄
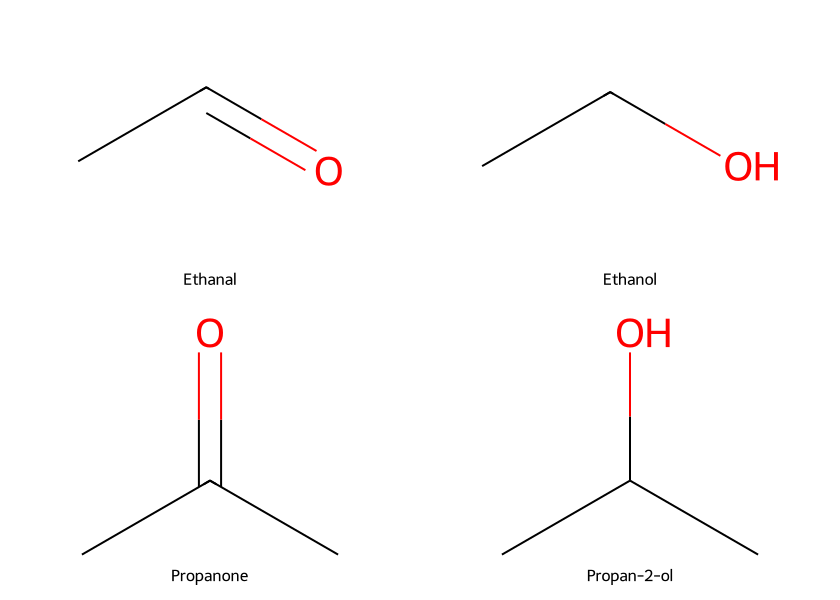
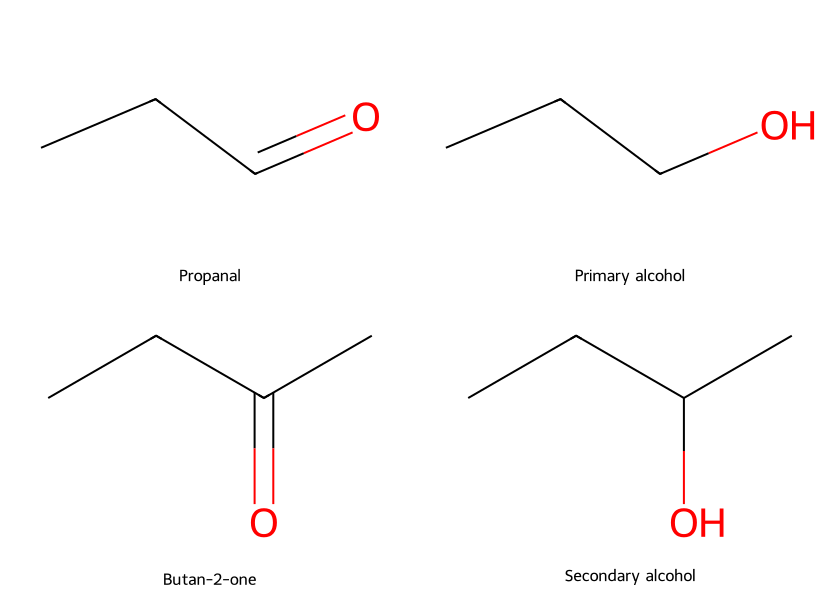
- NaBH₄ commonly reduces aldehydes to primary alcohols and ketones to secondary alcohols.
- LiAlH₄ also reduces aldehydes and ketones; it is stronger and requires careful anhydrous handling followed by workup.
- Hydride adds to the carbonyl carbon, then workup protonates the alkoxide.
| Transformation | Typical outcome |
|---|---|
| Aldehyde + NaBH₄ | Primary alcohol |
| Ketone + NaBH₄ | Secondary alcohol |
| Aldehyde + RMgX / workup | Secondary alcohol (formaldehyde → primary) |
| Ketone + RMgX / workup | Tertiary alcohol |
| Carbonyl + H₂O | Hydrate (geminal diol) |
| Carbonyl + 2 ROH / H⁺ | Acetal / ketal + water |
b. Grignard addition to carbonyl carbon
- Organomagnesium reagents (RMgX) behave as strongly nucleophilic carbon reagents.
- Addition to an aldehyde other than formaldehyde gives a secondary alcohol after aqueous or acidic workup; addition to a ketone gives a tertiary alcohol.
- Formaldehyde gives a primary alcohol after organomagnesium addition and workup.
- Grignard reagents are destroyed by water, alcohols, carboxylic acids, and other sufficiently acidic protons.
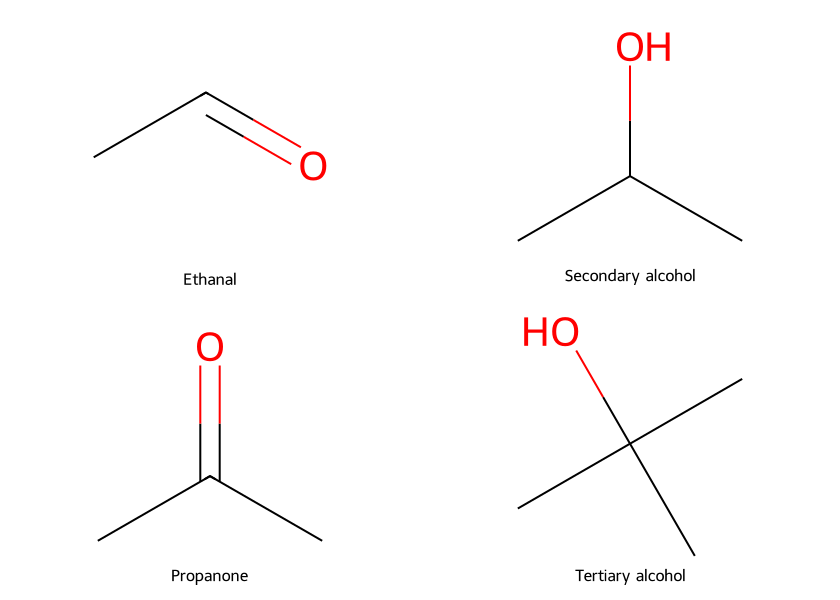
- Count the carbon atoms contributed by the incoming organometallic group.
- Organolithium reagents (RLi), like Grignard reagents, can make a new C–C bond and give an alcohol after protonation under compatible conditions.
c. Facial attack and stereochemical outcomes
- A planar achiral carbonyl can be attacked on either face. In an achiral environment, creating one new stereocenter commonly gives a racemic product.
d. Wittig alkene formation
- A phosphorus ylide replaces a carbonyl O with a carbon fragment to form C=C, with phosphine oxide as a byproduct.
- Disconnect the target alkene across its C=C to identify a carbonyl partner and the ylide carbon. E/Z selectivity depends on ylide type and conditions.
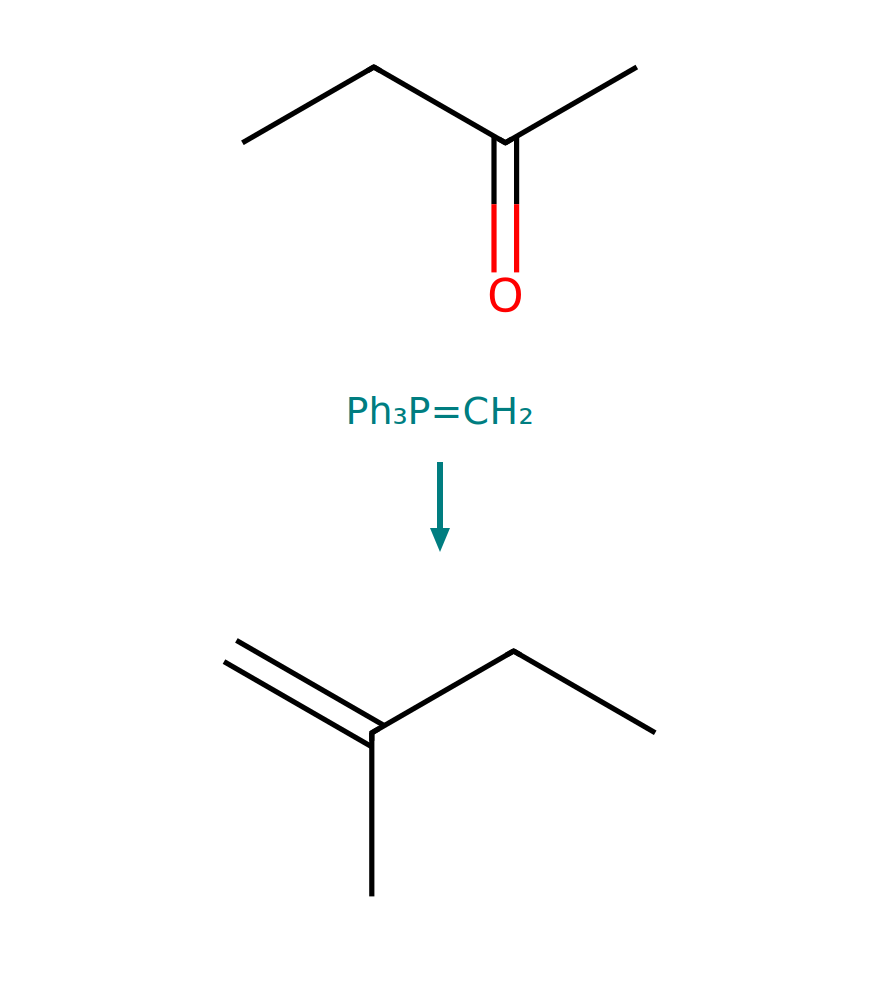
e. Carbonyl to methylene
- Clemmensen reduction uses strongly acidic Zn(Hg)/HCl conditions; Wolff–Kishner reduction uses hydrazine and strong base with heat. Both can remove carbonyl oxygen to give CH₂.
- Choose conditions compatible with the substrate. A thioacetal followed by suitable desulfurization is another carbonyl-to-methylene strategy noted in the source.
| Transformation | Typical method |
|---|---|
| C=O → C–OH | NaBH₄ or LiAlH₄; workup |
| C=O → C=C | Wittig reagent |
| C=O → CH₂, acidic | Zn(Hg), HCl |
| C=O → CH₂, basic | NH₂NH₂, base, heat |
f. Selective acyl chemistry
- An acid chloride can be reduced selectively to an aldehyde with a suitable bulky hydride, or converted to a ketone with an organocuprate. Nitrile addition by an organometallic reagent gives a ketone after hydrolysis.
- Ordinary Grignard addition to an ester usually proceeds beyond the ketone intermediate; a ketone synthesis requires a reagent/substrate combination that stops after the intended carbon addition.
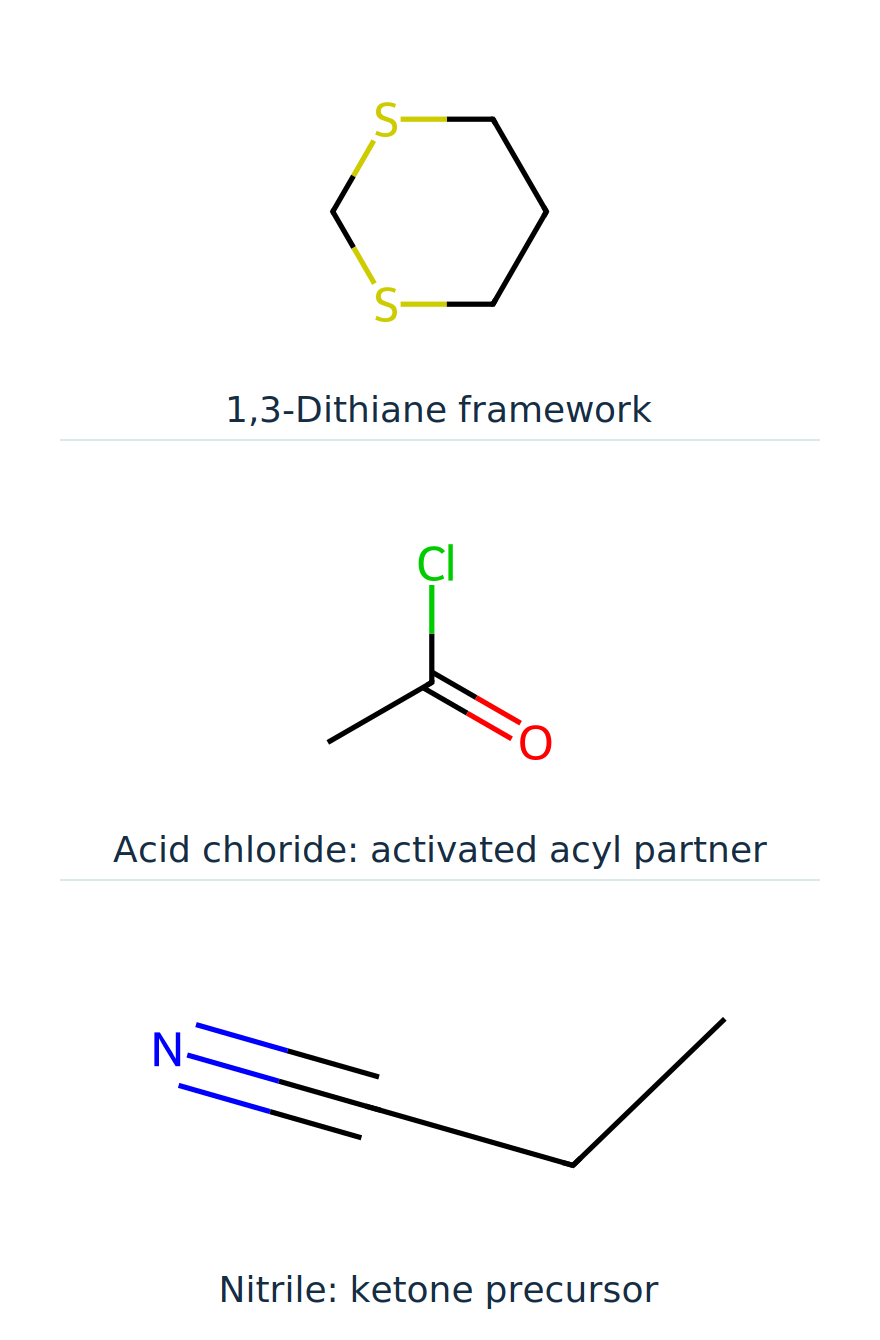
g. Dithiane strategy
- A 1,3-dithiane can act as a protected carbonyl and, after deprotonation at the carbon between sulfur atoms, as an acyl-anion equivalent. Alkylation at that carbon followed by appropriate hydrolytic deprotection reveals a carbonyl.
- This umpolung reverses the usual electrophilic role of carbonyl carbon. It is distinct from desulfurization, which removes sulfur to produce a methylene group.
D. Acetals and Imines
1. Reversible addition and condensation
a. Hemiacetal formation
- One equivalent of an alcohol may add to an aldehyde or ketone, producing a hemiacetal or hemiketal bearing both –OH and –OR on one carbon.
- Hemiacetals are important intermediates in acetal formation and occur in cyclic sugars.
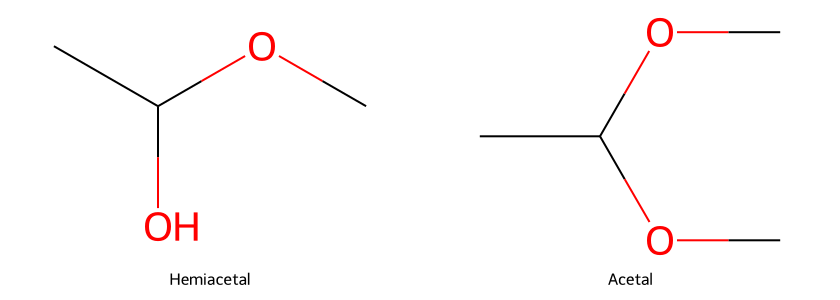
b. Acetal formation
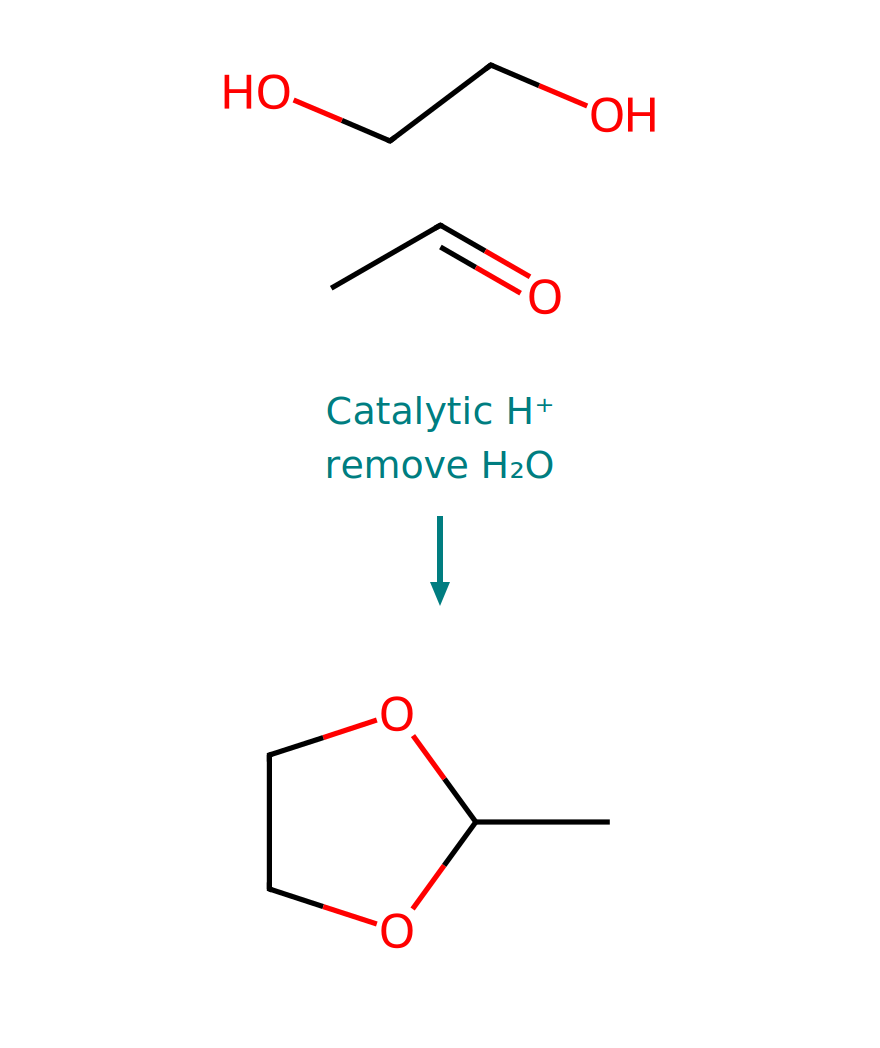
- Acid-catalyzed reaction with alcohol can yield an acetal, containing two –OR groups on the former carbonyl carbon.
- Excess alcohol or removal of water shifts the equilibrium toward acetal formation; aqueous acid promotes hydrolysis back to the carbonyl compound.
- Acetals can protect a carbonyl group during selected synthetic transformations.
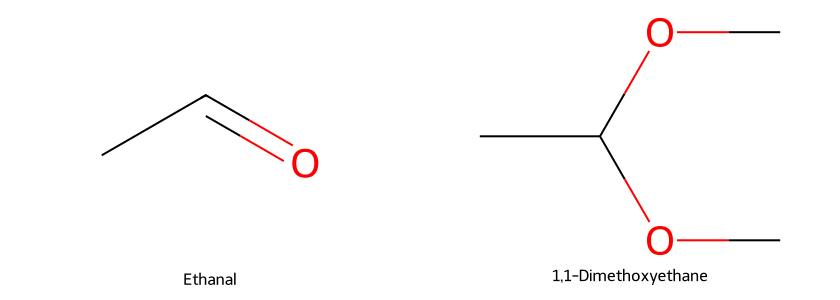
- The acetal center no longer contains a C=O bond.
- Suitable diols can form cyclic acetals.
c. Protecting-group logic
- Acetals generally tolerate many basic nucleophilic conditions but hydrolyze under aqueous acid. They can protect a carbonyl during an incompatible step.
- Protection and deprotection add steps and must be compatible with the other functional groups in the molecule.
d. Condensation
- A primary amine can form an imine after addition, proton transfer and loss of water. Mild acid catalysis is useful, while strong acid can suppress nucleophilicity by protonating the amine.
- A secondary amine can form an enamine when the carbonyl partner has an appropriate α-H. The product contains a C=C adjacent to N.
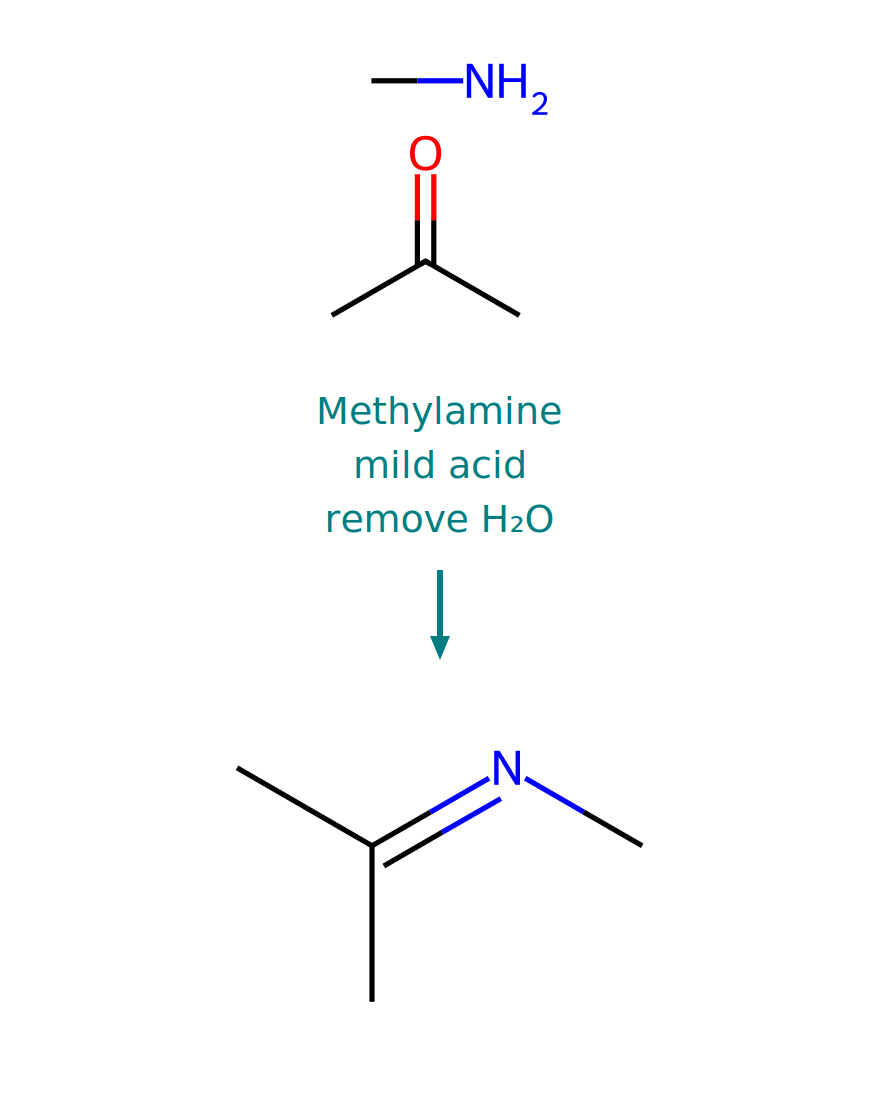
- Imine formation proceeds through a carbinolamine intermediate before loss of water.

e. Related derivatives
- Hydroxylamine forms oximes; hydrazine derivatives form hydrazones. These are condensation products rather than simple alcohol products.
- Reductive amination reduces an imine/iminium intermediate to an amine and is developed with nitrogen chemistry.
E. Enolates and Aldol Reactions
1. Tautomerism, alpha chemistry and carbon–carbon formation
a. Interconversion
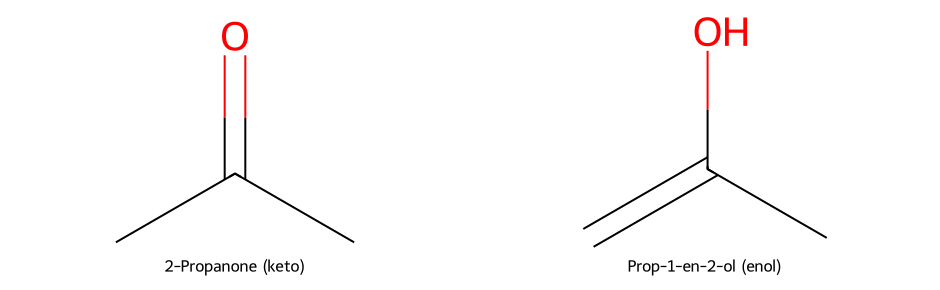
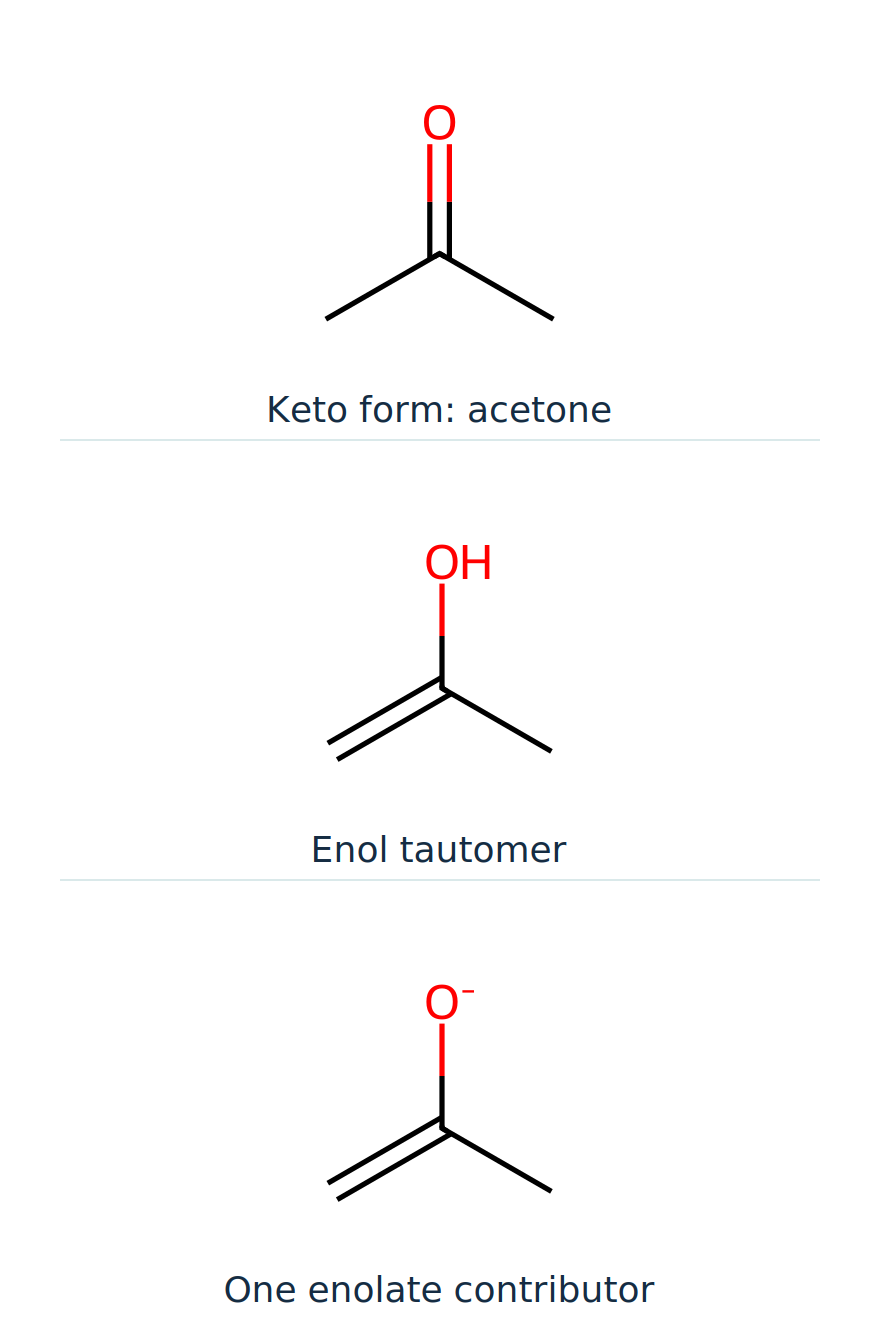
- A carbonyl compound with an alpha hydrogen may interconvert with its enol tautomer.
- Keto and enol tautomers differ in proton location and double-bond placement; they are distinct structures, not resonance contributors.
- For many simple aldehydes and ketones, the keto form is favored at equilibrium.
b. Acid- and base-mediated pathways
- Acid catalysis involves carbonyl oxygen protonation followed by removal of an alpha proton.
- Base catalysis involves alpha deprotonation to an enolate followed by protonation at oxygen to produce an enol.
c. Enolate stabilization
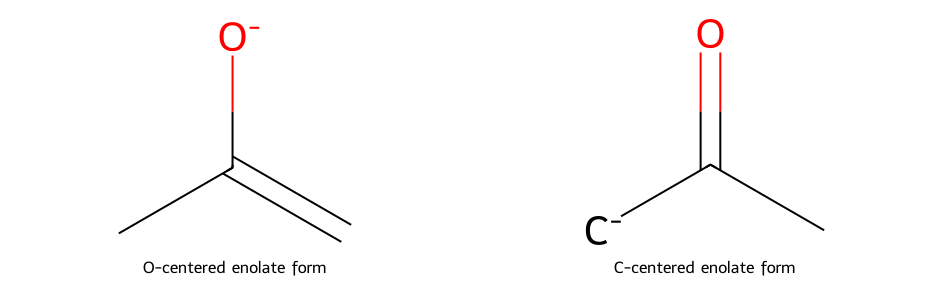
- An enolate delocalizes negative charge between α-carbon and oxygen. Reactions at either site can occur depending on electrophile and conditions.
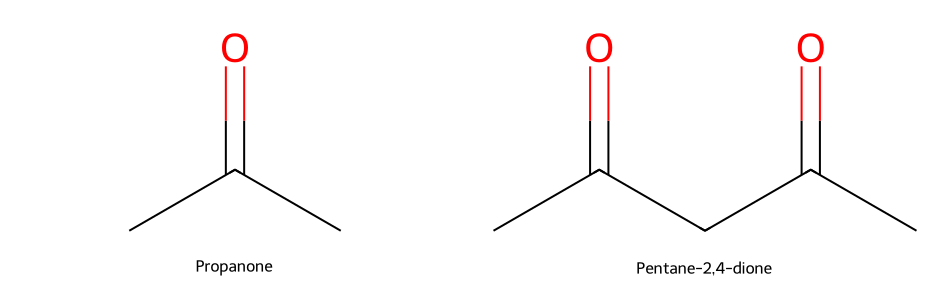
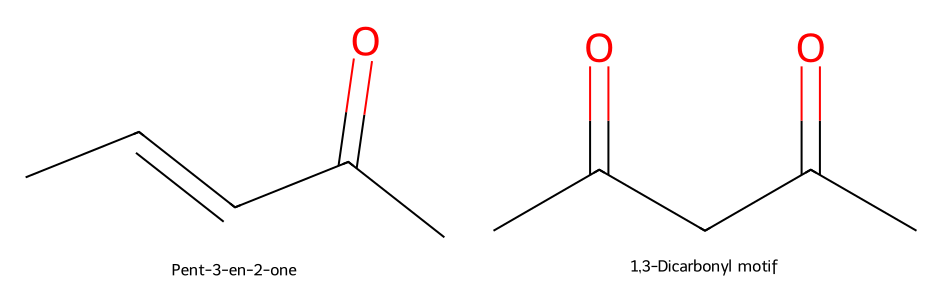
- Typical ketone α-H atoms are much more acidic than alkane H atoms. A β-dicarbonyl compound is more acidic still because its conjugate base can interact with two carbonyl groups.
| Acidic site | Typical approximate pKa |
|---|---|
| β-Diketone central C–H | about 9 |
| β-Keto ester central C–H | about 11 |
| Ketone α-C–H | about 19–21 |
| Ester α-C–H | about 25 |
- Enolate resonance contributors retain atom connectivity while delocalizing charge between oxygen and the α-carbon.
d. Enolate alkylation at the alpha carbon
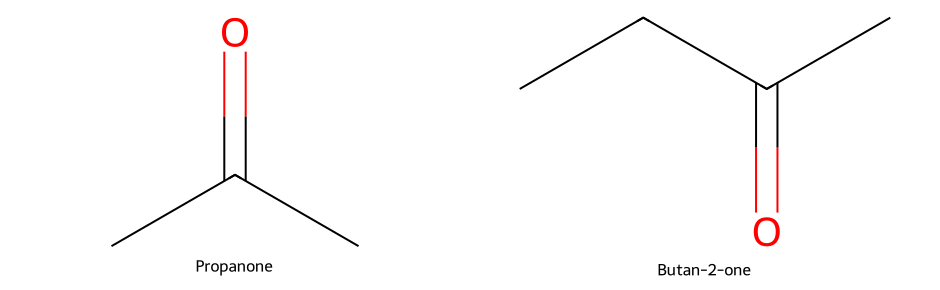
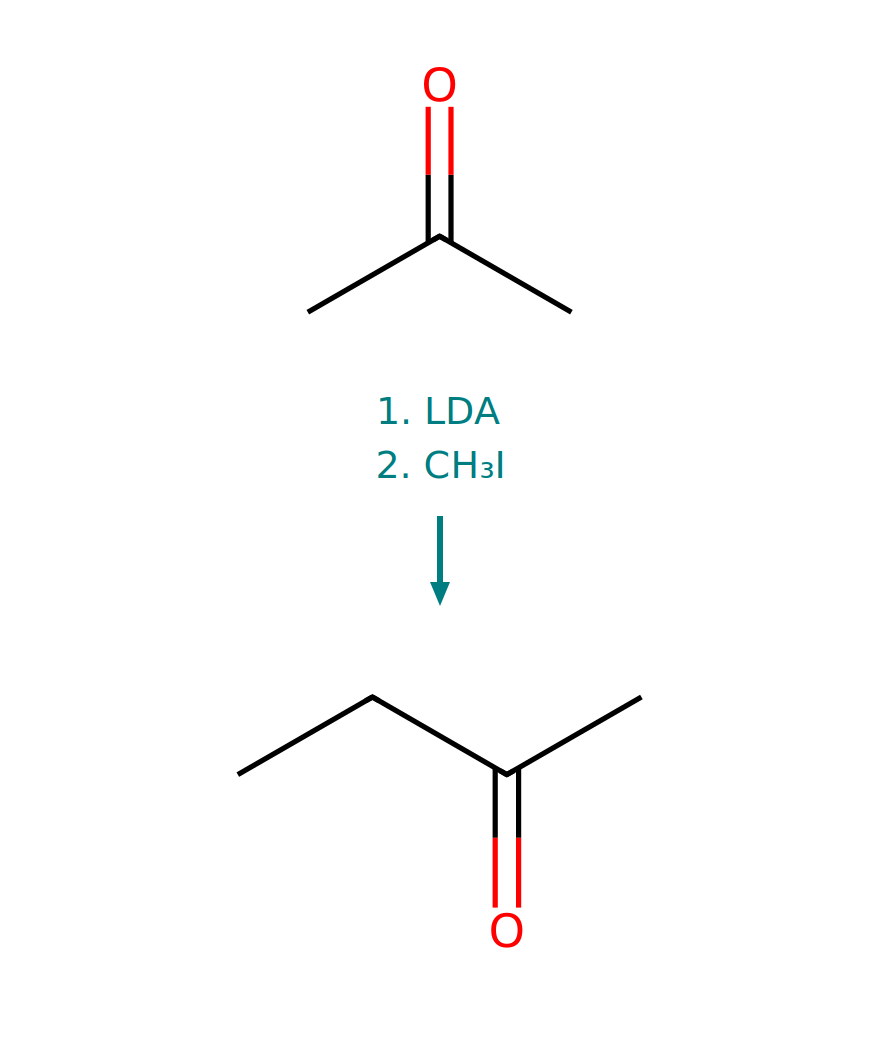
- A carbon-centered enolate can react with a suitable primary alkyl halide in an SN2 process.
- The reaction forms a new C–C bond at the alpha position.
- Steric hindrance and competing elimination can limit reactions with secondary or tertiary electrophiles.
- A sufficiently strong base is needed to generate an enolate; a relatively nonnucleophilic base such as LDA is useful for substantial deprotonation under controlled conditions.
e. Regioselectivity
- An unsymmetrical carbonyl can form different enolates. Kinetic versus thermodynamic enolate selection depends on base, temperature, solvent and equilibration.
- Repeated alkylation and competing O-alkylation can occur. Plan both the selected α-site and the electrophile, rather than treating all enolates as interchangeable.
f. Acid versus base
- Acidic conditions often give controlled monohalogenation through the enol. Under basic conditions, each installed halogen can increase the acidity of remaining α-H atoms and encourage further halogenation.
- For carboxylic acids with an α-H, Hell–Volhard–Zelinsky conditions enable α-halogenation through an activated acyl intermediate, followed by hydrolysis.

g. Haloform reaction
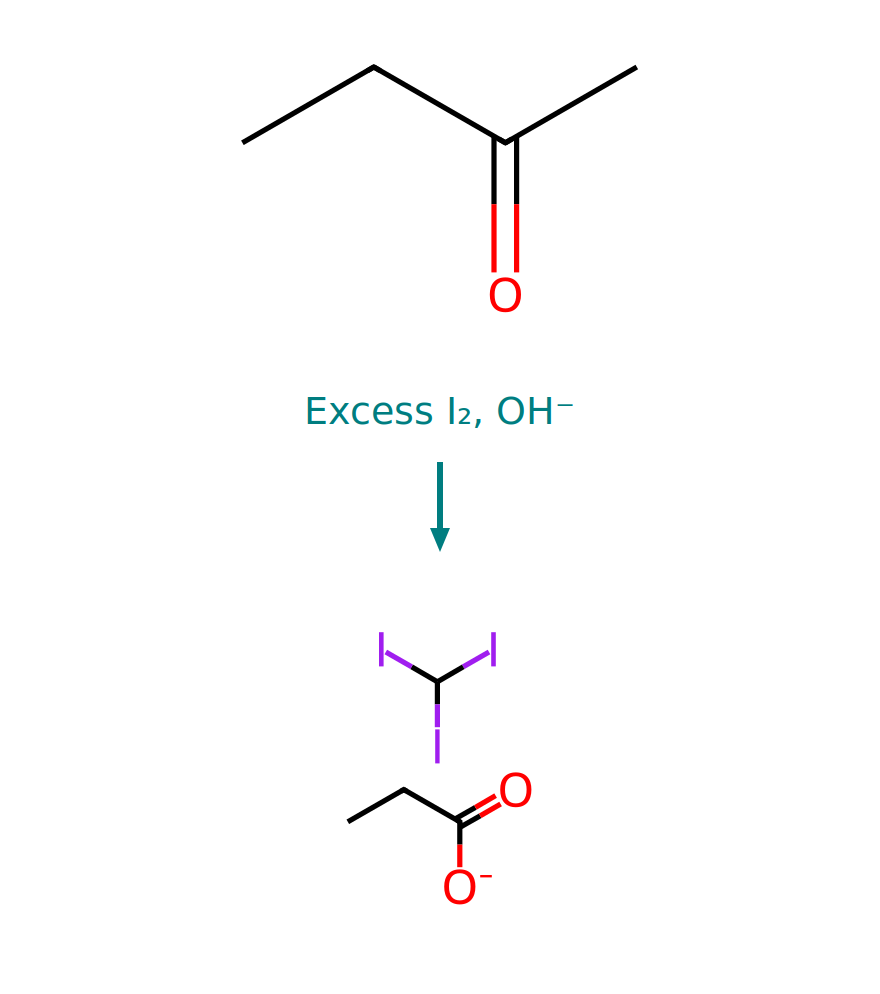
- A methyl ketone can undergo exhaustive α-halogenation and cleavage under suitable X₂/OH⁻ conditions to yield carboxylate and CHX₃.
- The iodoform reaction can also be positive for alcohols oxidized under the conditions to suitable methyl ketones, as well as ethanol/acetaldehyde. Product interpretation depends on the starting functional group.
h. Addition step
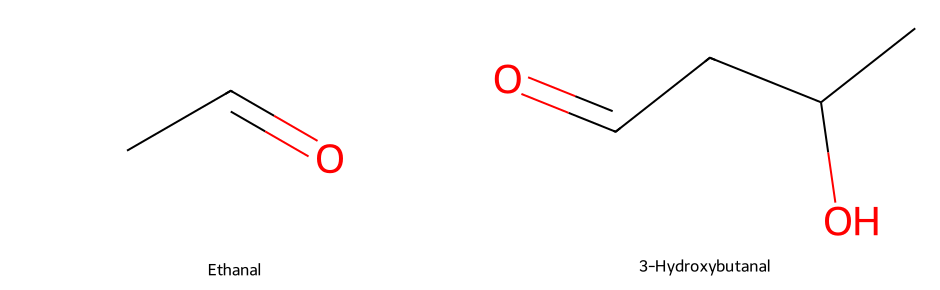
- An enolate adds to another aldehyde or ketone. Protonation gives a β-hydroxy carbonyl product: the electrophile carbonyl becomes OH while the donor carbonyl remains.
- The donor needs an enolizable α-site; the acceptor must have an accessible electrophilic carbonyl.

- An enol-derived carbon nucleophile can also participate under suitable conditions; trace the new bond from the donor α-carbon to the acceptor carbonyl carbon.
i. Dehydration
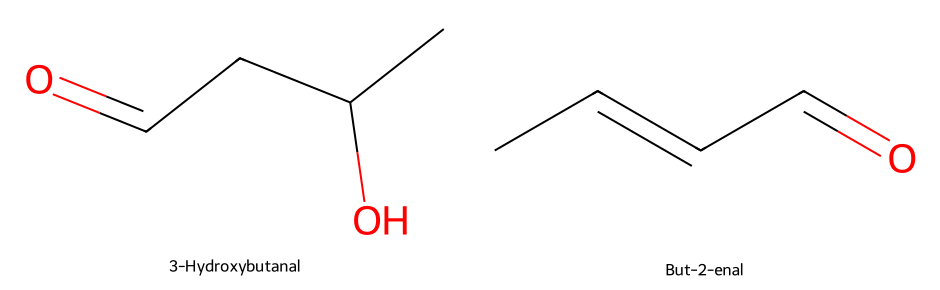
- On heating or under suitable acid/base conditions, a beta-hydroxy carbonyl compound can lose water.
- The resulting alpha,beta-unsaturated carbonyl system is conjugated.
- Aldol addition and aldol condensation are different stages; dehydration is not automatic in every experiment.
- Under basic conditions, suitable aldol dehydration commonly follows an E1cb-type sequence.
j. Selectivity
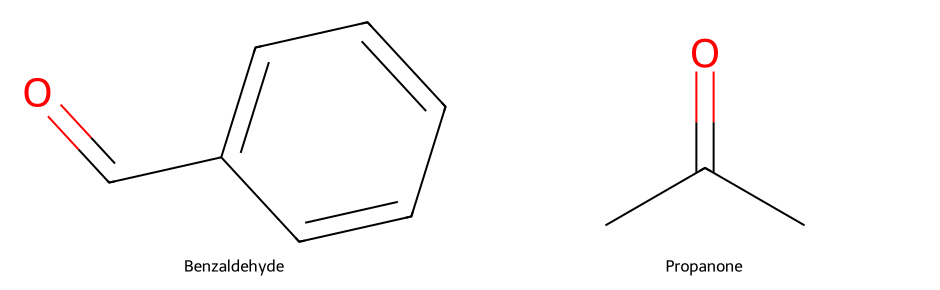
- When two different enolizable carbonyl partners are mixed, multiple aldol products may be possible.
- A carbonyl partner lacking alpha hydrogens, such as benzaldehyde, cannot serve as a normal enolate donor.
- Preforming a specific enolate can improve selectivity.
k. Ring formation
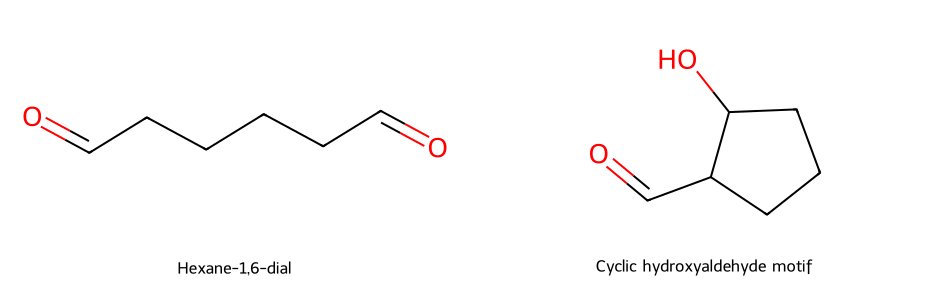
- A molecule containing two suitably spaced carbonyl groups may undergo intramolecular aldol chemistry.
- Formation of common five- or six-membered rings is often favored when accessible.
l. 1,2 versus 1,4
- An α,β-unsaturated carbonyl has electrophilic sites at carbonyl carbon and β-carbon. Direct 1,2-addition attacks C=O; conjugate 1,4-addition attacks β-carbon.
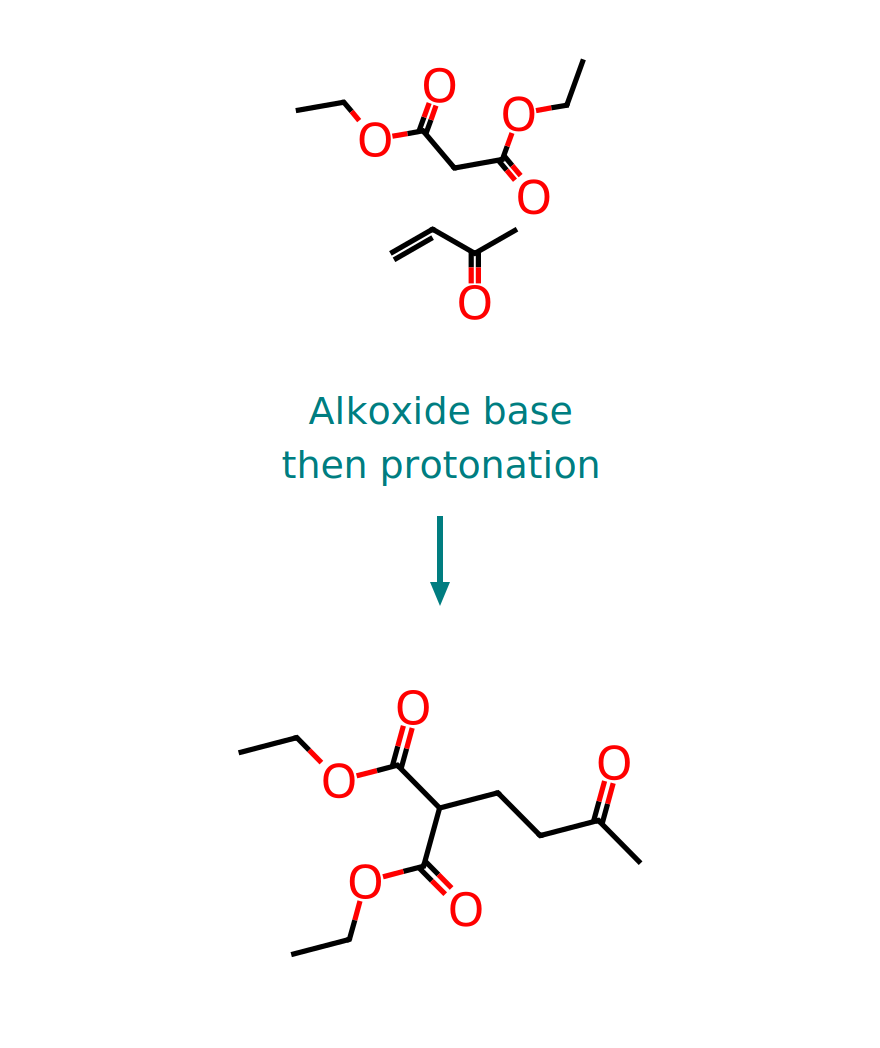
- Stabilized carbon nucleophiles, such as many β-dicarbonyl enolates, commonly participate in Michael addition.
m. Product connectivity
- After conjugate attack and proton transfer, the acceptor carbonyl is retained. A carbonyl-containing donor can produce a 1,5-dicarbonyl relationship.
- A synthesis plan should track the new donor-α to acceptor-β bond and account for every carbon before choosing reagents.