
CH 5. Alkenes & Alkynes
A. Structures and Stability
1. Unsaturation, reactive species and reaction energy
a. Structures, molecular formulas and orbital geometry
- Alkenes contain at least one carbon–carbon double bond; alkynes contain at least one carbon–carbon triple bond.
- An acyclic monoalkene has molecular formula CₙH₂ₙ; an acyclic monoalkyne has CₙH₂ₙ₋₂. These expressions do not apply to every ring-containing or multiply unsaturated structure.
- Unsaturation changes orbital geometry and enables reactions at the π bond.
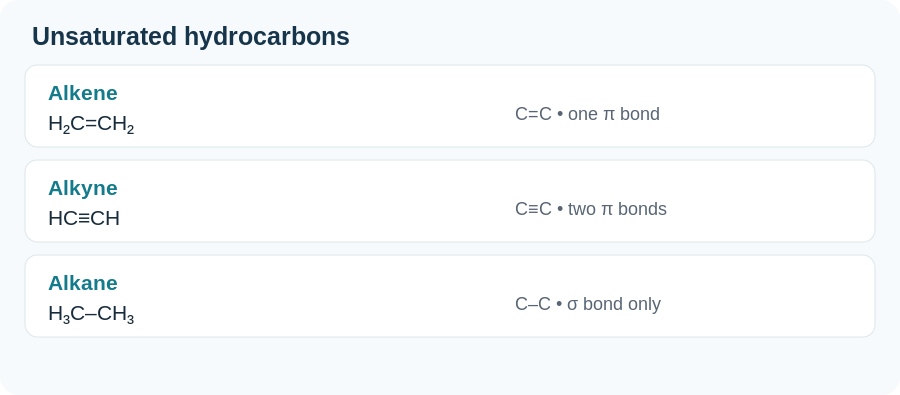
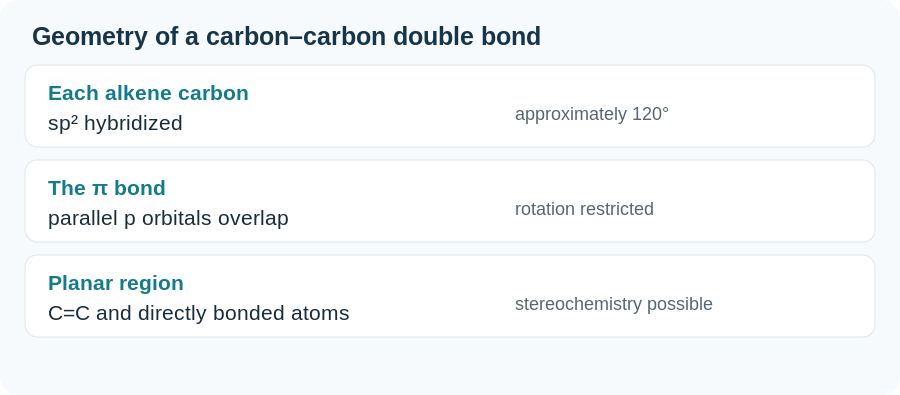
- Alkene carbons are approximately sp² and trigonal planar; alkyne carbons are sp and linear. A multiple bond contains one σ bond plus one or two π bonds.
- Twisting the ends of C=C disrupts π overlap.
b. Selecting the Parent
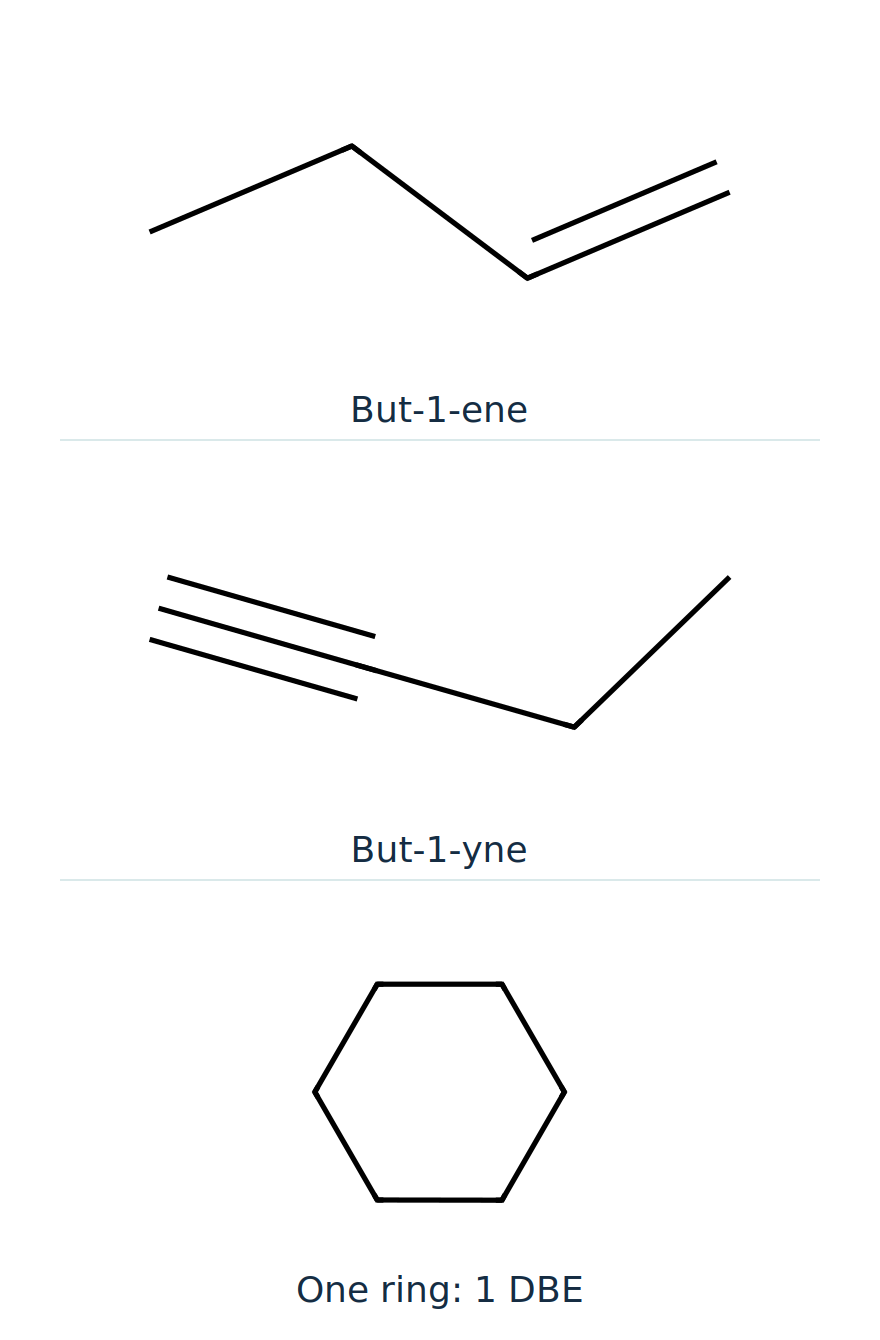
- Choose the parent framework containing the multiple bond and number it to assign appropriate low locants to unsaturation.
- Use -ene and -yne endings with locants for the position of the double or triple bond.

c. Multiple Unsaturations
- Two double bonds are described by the suffix -diene; three by -triene.
- A molecule containing both C=C and C≡C bonds uses an en-yne name and appropriate locants.
d. Addition and Elimination
- In addition, atoms or groups add across a multiple bond and its π bond is converted to σ bonds.
- In elimination, the substrate loses atoms or groups and a π bond can form.

e. Substitution and Rearrangement
- Substitution replaces a leaving group with another group.
- Rearrangement changes the connectivity of atoms within a molecular framework without necessarily changing molecular formula.
f. Degree of unsaturation
- For a usual neutral closed-shell formula containing C, H, N and halogens, DBE = (2C + 2 + N − H − X)/2. Oxygen and divalent sulfur do not enter this count.
- A ring or double bond contributes one DBE; a triple bond contributes two. DBE constrains possibilities but does not determine a unique structure.
| Feature | DBE contribution |
|---|---|
| Ring | 1 |
| C=C or C=O | 1 |
| C≡C or C≡N | 2 |
| Benzene ring | 4 |
g. Pair-electron reactions
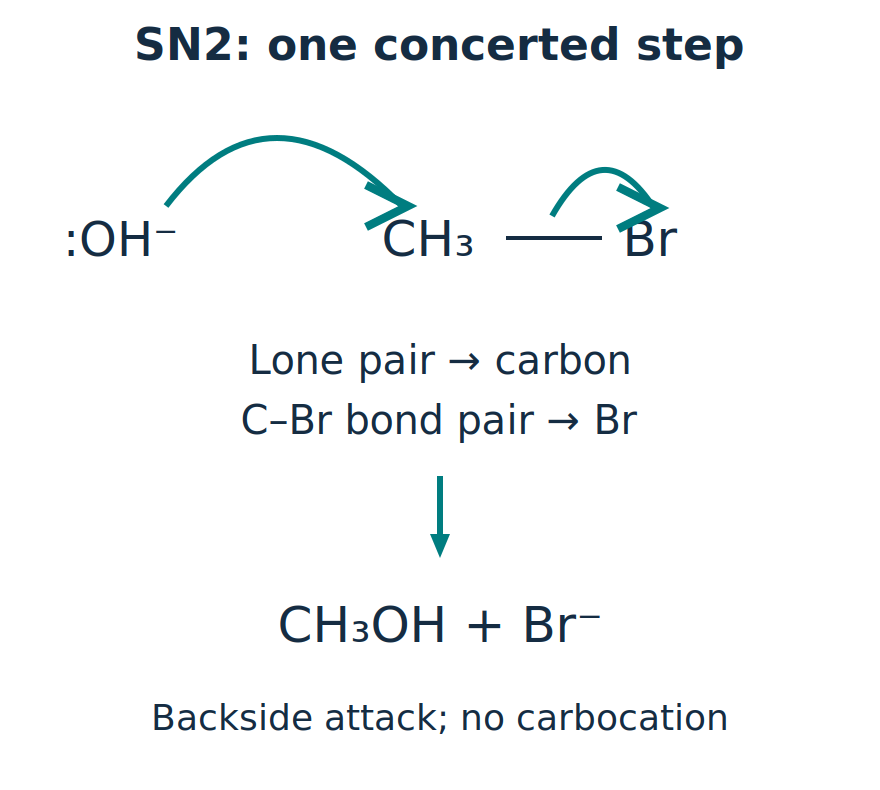
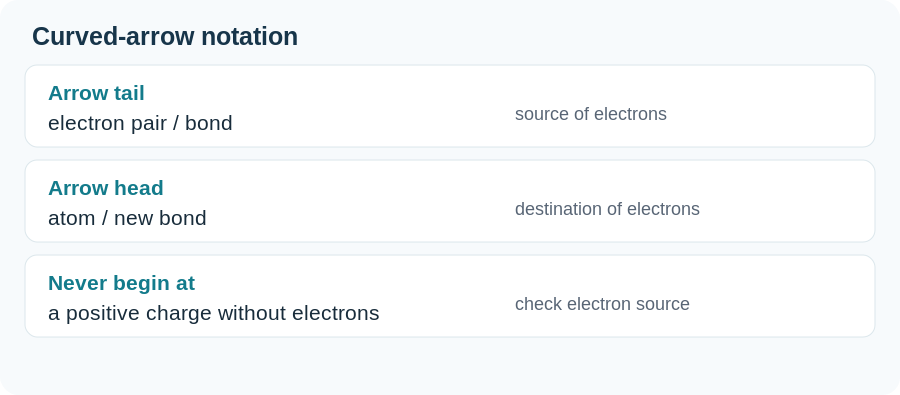
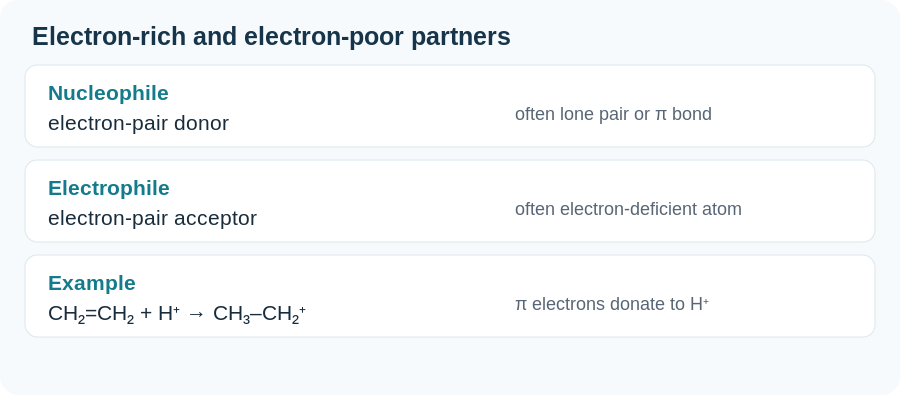
- A nucleophile donates an electron pair; an electrophile accepts one. A lone pair, negative charge or pi bond can provide the electron source.
- A curved arrow begins at electrons and ends where a bond or lone pair forms. It never begins at an isolated positive charge.
- An alkene π bond can act as the electron-rich partner in electrophilic addition.
h. Single-electron reactions
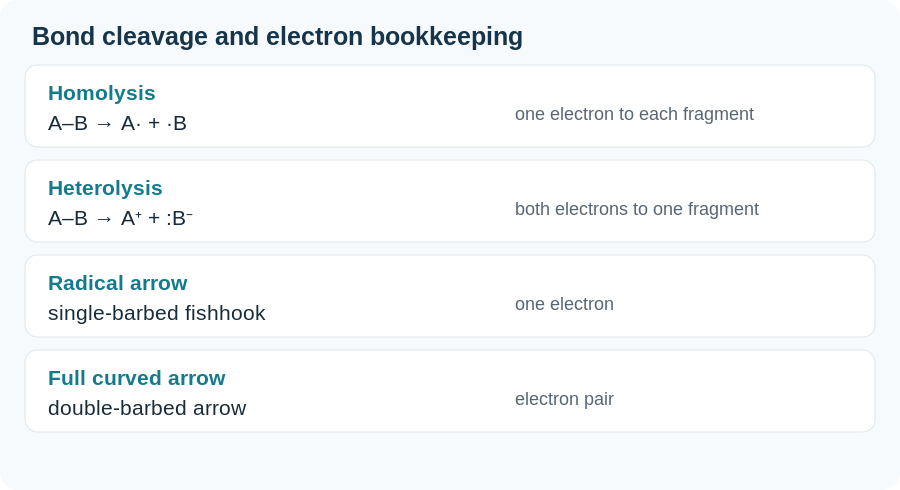
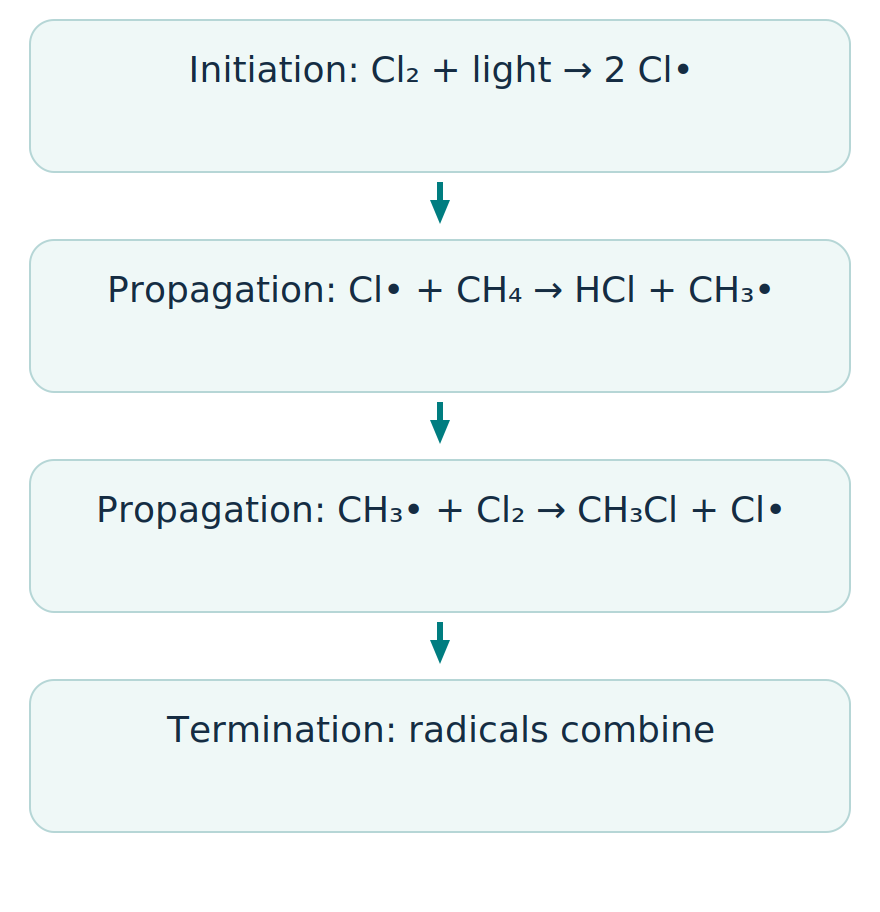
- Homolytic bond cleavage forms radicals. Radical mechanisms use fishhook arrows and often form chains with initiation, propagation and termination.
- In heterolysis, the entire bonding pair remains on one fragment. The resulting ions have charges that must balance the starting material.
i. Energy diagrams
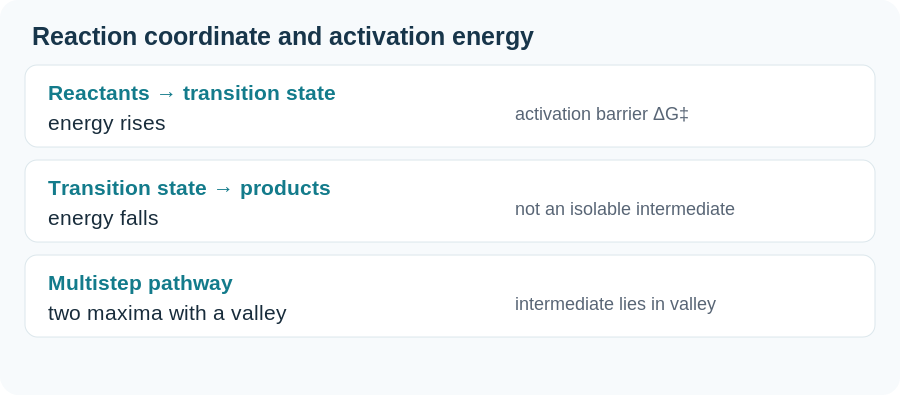
- A transition state is an energy maximum along the reaction coordinate; an intermediate is a local minimum between maxima.
- The activation free energy controls rate. The free-energy difference between reactants and products controls equilibrium. A catalyst changes the pathway and rate but not the equilibrium constant.
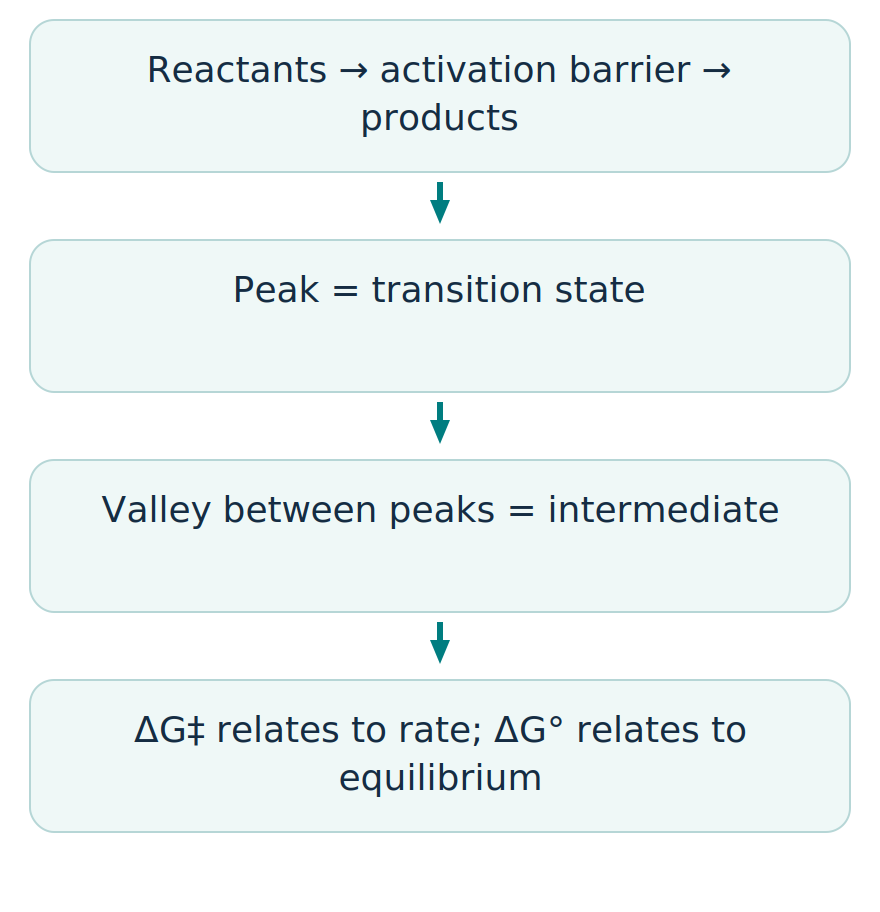
- A transition state is not an isolable species.
j. Temperature and the Hammond idea
- Increasing temperature usually increases reaction rate. For closely related pathways, a higher barrier generally gives a slower reaction at the same temperature.
- A transition state tends to resemble the adjacent species closest to it in energy. Late, product-like radical abstraction transition states often show greater selectivity.
| Quantity | Meaning |
|---|---|
| ΔG‡ | Activation free energy |
| ΔG° | Overall free-energy change |
| ΔH ≈ ΣD(broken) − ΣD(formed) | Approximate bond-enthalpy estimate |
k. Carbocations and radicals
- Simple carbocations are usually planar and electron deficient; alkyl substitution often stabilizes them by hyperconjugation and induction. Allylic and benzylic resonance can dominate the alkyl trend.
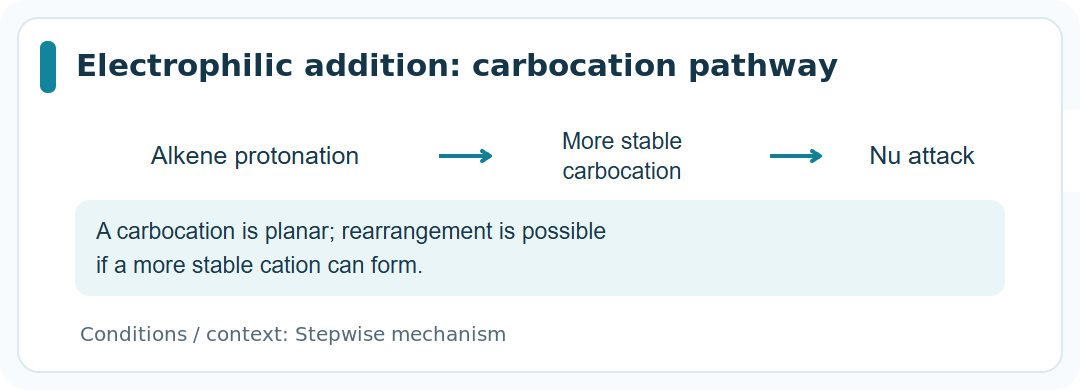
- A carbocation can undergo a 1,2-hydride or alkyl shift if the resulting pathway is favorable. Check connectivity as well as the final functional group.

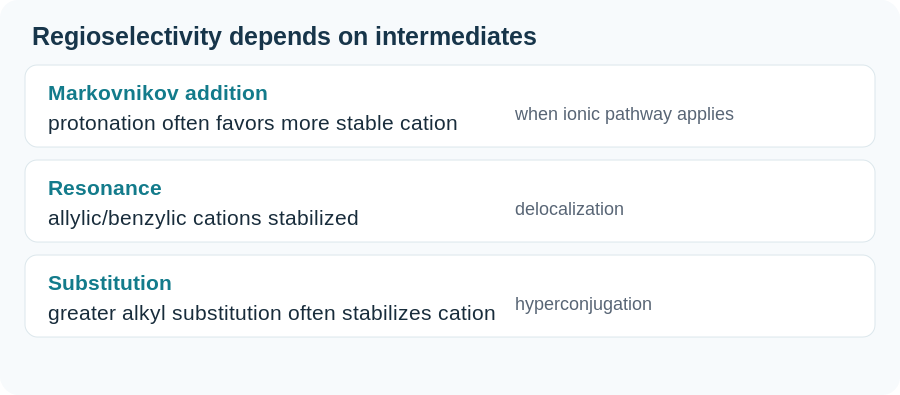
- Protonation consumes the alkene π bond. For simple alkyl carbocations, tertiary > secondary > primary is a starting stability trend; resonance can override it. A planar carbocation can be attacked on either face and may give a stereoisomer mixture.
l. Carbanions and carbenes
- Carbanion stability depends strongly on resonance, electronegative substituents, hybridization and solvent. A simple alkyl trend cannot be applied across unlike structures.
- A carbene is a neutral divalent carbon species. Carbene and carbenoid additions can convert an alkene into a cyclopropane.
m. Chain sequence
- Light can cleave X–X to initiate radicals. A halogen radical abstracts H, then the carbon radical reacts with X₂ to form product and regenerate X•.
- Termination combines radicals without regenerating a chain carrier. Product ratios depend on both the number of available hydrogens and their relative reactivities.
n. Chlorination and bromination
- Bromination is usually more selective for the site forming the more stable radical; chlorination is less selective.
- Radical halogenation can yield mixtures and is not a universal way to obtain a single alkyl halide. Allylic bromination with NBS is treated with conjugated systems.
o. Comparing heats of hydrogenation
- Compare heats of hydrogenation only for genuinely comparable molecules and products.
B. Electrophilic Addition
1. Ionic and radical addition pathways
a. Ionic HX addition and carbocation regioselectivity
- The alkene π bond donates an electron pair to H from HX; halide then captures the carbocation to form C–X. For an unsymmetrical alkene, the favored pathway commonly forms the more stable carbocation and gives Markovnikov orientation.
- Hydride or alkyl shifts can change the carbon skeleton before capture. This stepwise carbocation pathway is generally not stereospecific.
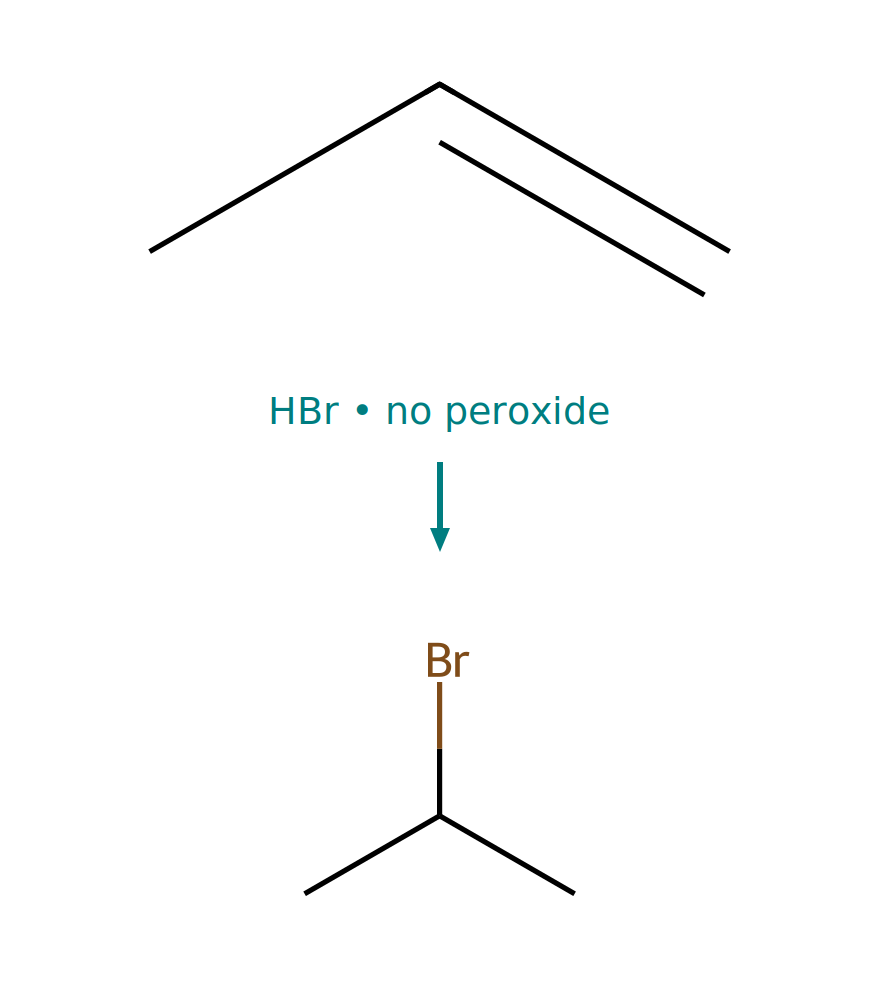

b. HBr addition to ethene: a concrete ionic example
- In a simplified ionic addition of HBr to ethene, the π bond forms a C–H bond as the H–Br bond cleaves heterolytically.
- Bromide subsequently bonds to the carbocationic carbon, forming bromoethane.
- More complex substrates may rearrange or display other selectivity depending on reagents and conditions.
c. HBr with peroxides
- Under appropriate radical conditions, Br adds so that the more stable carbon radical forms; H abstraction from HBr completes anti-Markovnikov addition.
- The useful peroxide effect is characteristic of HBr. Do not extend it automatically to HCl or HI.
| Conditions | Orientation | Intermediate |
|---|---|---|
| HX, ordinary ionic conditions | Usually Markovnikov | Carbocation |
| HBr, ROOR, radical conditions | Anti-Markovnikov | Carbon radical |
d. Dihalide formation
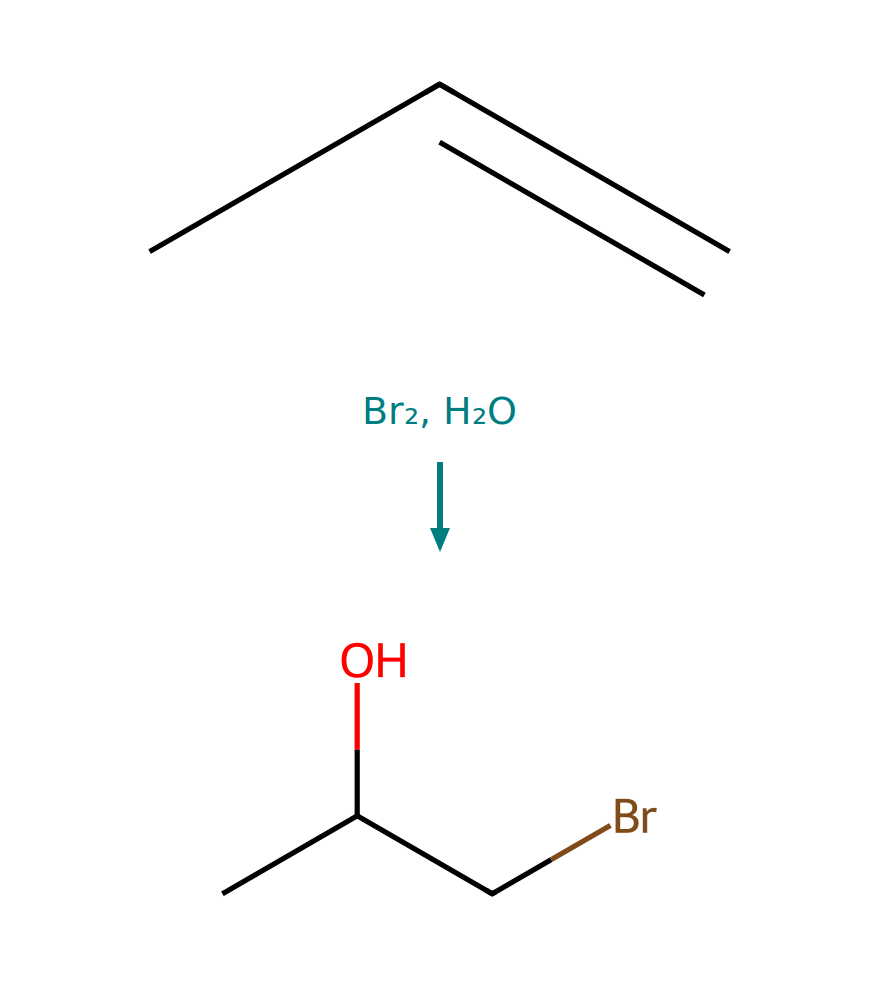
- Br₂ or Cl₂ adds through a bridged halonium ion. Backside opening commonly gives anti addition across the former double bond.
- A halonium pathway avoids ordinary hydride and alkyl shifts. The substrate geometry controls the stereochemical relationship of products.
- For cyclic substrates, use the starting geometry and facial accessibility when assigning stereochemical relationships.
- In an achiral medium, an achiral alkene can give a racemic pair when the addition creates new stereocenters.
e. Water as nucleophile
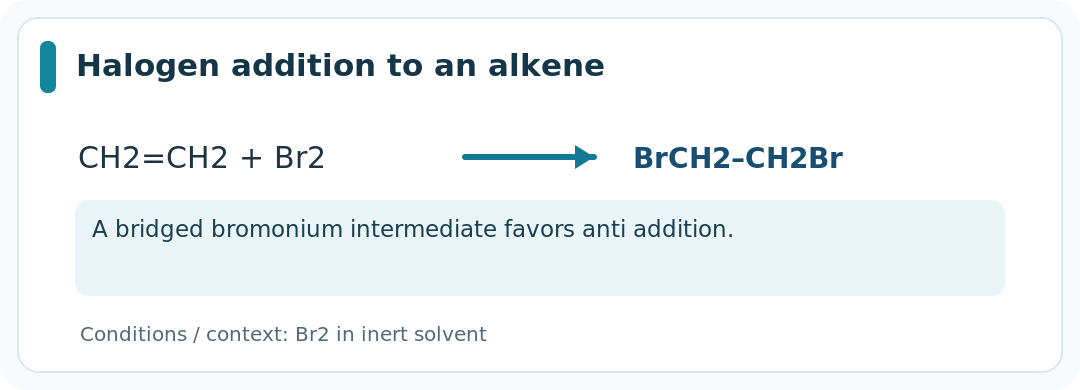
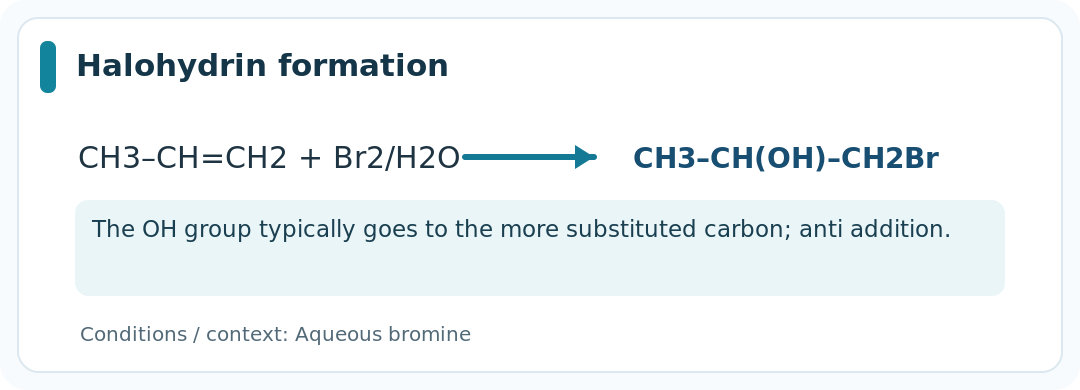
- With Br₂/H₂O, water opens the bromonium ion, followed by deprotonation. OH normally appears at the more substituted carbon and Br at the other.
- The OH and halogen are anti. Formation of an enantiomeric pair depends on substrate symmetry and whether a chiral influence is present.
f. Chain growth from alkene monomers
- Alkene molecules can combine into long chains by converting π bonds into new σ bonds.
- A radical polymerization proceeds through initiation, chain propagation, and termination.
- The polymer repeat unit retains the carbon framework of the monomer; polyethylene has the repeat unit –CH2–CH2–.
- The average polymer length and properties depend on the initiation, propagation, transfer, and termination processes.

C. Hydration and Hydroboration
1. Regioselectivity and hydration mechanisms
a. Addition of water
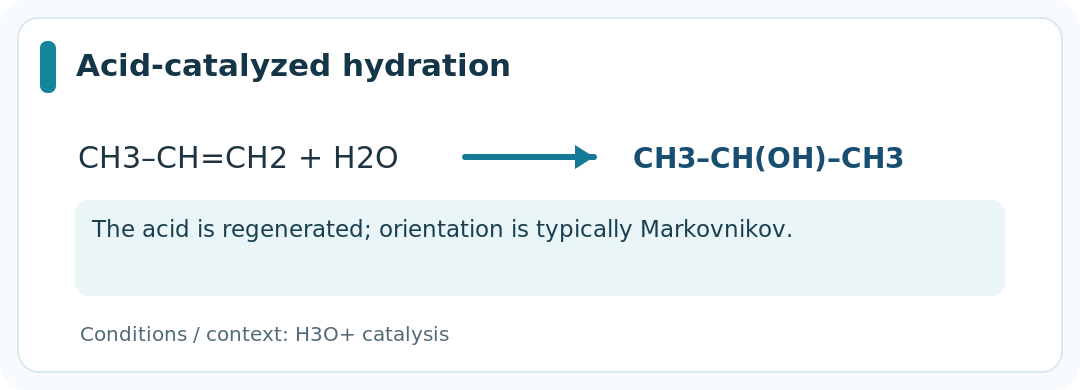
- Dilute aqueous acid hydrates alkenes by protonation, water attack, and deprotonation.
- Regioselectivity generally corresponds to Markovnikov addition, with OH at the more substituted former alkene carbon.
- Since a carbocation may form, hydride or alkyl shifts can compete in susceptible substrates.
- The hydration/dehydration equilibrium depends on conditions and substrate.
b. Acid and oxymercuration
- Dilute aqueous acid gives Markovnikov hydration through protonation, water attack and deprotonation; rearrangements may occur.
- Oxymercuration–demercuration uses Hg(OAc)₂/H₂O then NaBH₄ to give Markovnikov alcohols without typical free-carbocation rearrangements. Overall stereochemistry is not reliably a simple anti rule.
c. Hydroboration–oxidation: syn anti-Markovnikov hydration
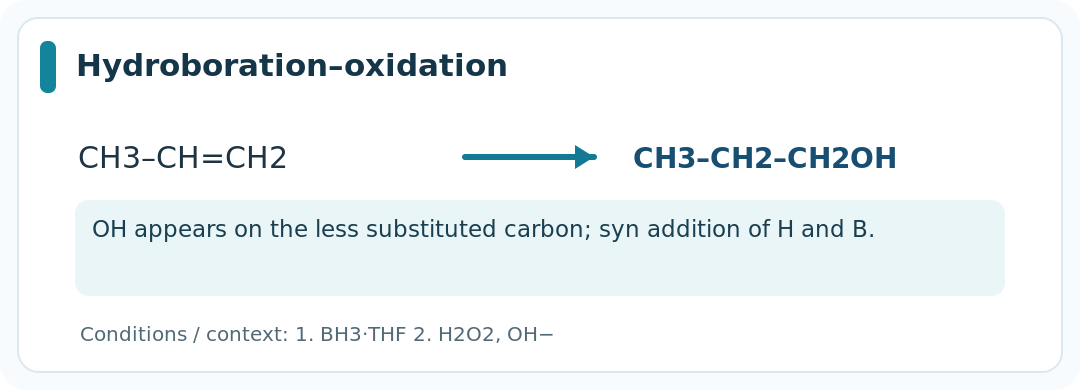
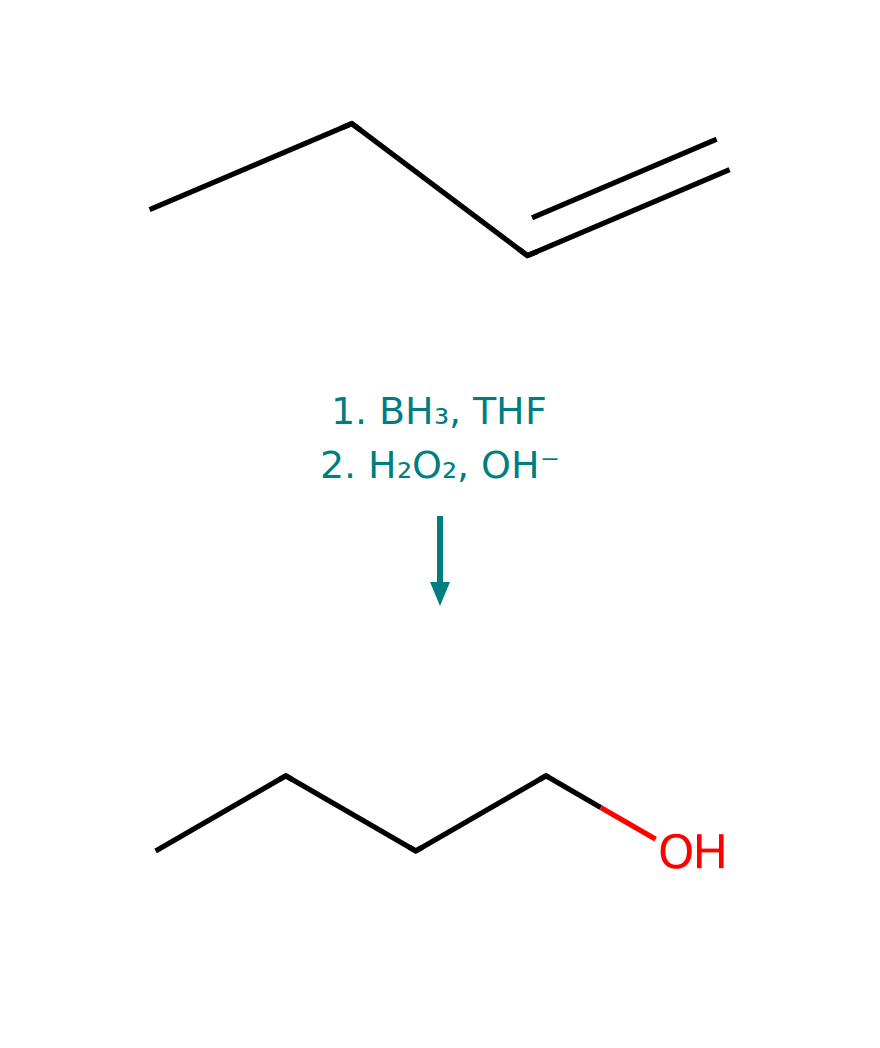
- BH₃, commonly supplied in THF, adds H and B across C=C in a concerted syn process. B generally attaches to the less substituted carbon.
- Subsequent H₂O₂/OH⁻ oxidation replaces C–B with C–OH with retention at that carbon. The overall hydration is syn and anti-Markovnikov, without a freely rearranging carbocation.
| Method | OH position | Rearrangement |
|---|---|---|
| H₃O⁺ | More substituted | Possible |
| Hg(OAc)₂/H₂O; NaBH₄ | More substituted | Normally absent |
| BH₃; H₂O₂/OH⁻ | Less substituted | Absent |
- Track the carbon skeleton and both possible faces when a new stereocenter can form.
D. Oxidation and Reduction
1. Hydrogenation, oxidation and alkyne transformations
a. Reduction of carbon–carbon double bonds
- H2 in the presence of Pt, Pd, or Ni converts many C=C bonds into C–C single bonds.
- Hydrogen is transferred from the metal surface and the net addition to an alkene is syn.
- Catalytic hydrogenation decreases unsaturation without changing the number of carbon atoms.
- When a cyclic alkene is reduced, approach to the two faces may be influenced by steric accessibility.
b. Epoxides, diols, and oxidative cleavage
- Peroxyacids such as mCPBA convert alkenes to three-membered epoxide rings in a concerted oxygen-transfer step.
- Syn dihydroxylation can produce a vicinal diol with two OH groups added to the same alkene face.
- Ozonolysis cleaves the C=C bond to carbonyl compounds under reductive workup.
- The oxidative workup of ozonolysis can convert aldehyde products into carboxylic acids.
c. Terminal alkynes as carbon nucleophiles
- Terminal alkynes are more acidic than comparable alkenes or alkanes because the conjugate base has negative charge in an sp orbital.
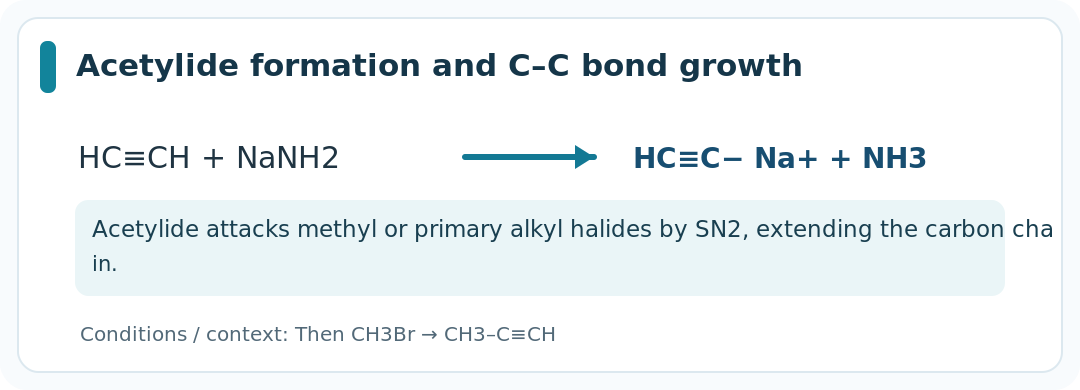
- A sufficiently strong base such as NaNH2 converts RC≡CH into an acetylide anion, RC≡C−.
- Acetylide anions form C–C bonds by SN2 attack on methyl and many primary alkyl halides.
- Secondary and tertiary halides are poor substrates for acetylide SN2 alkylation because competing elimination or steric obstruction dominates.
- Alkylation extends the carbon framework; check how many new carbon atoms are added.
| Reagent | Net result | Regiochemistry |
|---|---|---|
| HBr | Alkyl bromide | Often Markovnikov |
| H3O+ | Alcohol | Markovnikov |
| BH3; H2O2/OH− | Alcohol | Anti-Markovnikov |
| Br2 | Vicinal dibromide | Anti addition |
| Br2/H2O | Halohydrin | OH at more substituted carbon |
| H2/Pd | Alkane | Syn reduction |
| Reagent | Net result | Selectivity |
|---|---|---|
| H2/Lindlar | cis alkene | Syn partial reduction |
| Na/NH3(l) | trans alkene | Anti partial reduction |
| HgSO4/H2SO4/H2O | Ketone | Terminal (except acetylene) → methyl ketone; acetylene → ethanal |
| 9-BBN; H2O2/OH− | Aldehyde | Terminal → aldehyde |
| NaNH2; primary R–X | Higher alkyne | C–C bond formation |
d. Carbene and carbenoid cyclopropanation
- A suitable carbene or carbenoid can add across an alkene to form a cyclopropane. Simmons–Smith conditions use CH₂I₂ and a zinc reagent and retain the alkene’s relative stereochemistry.
e. Epoxide and diol formation
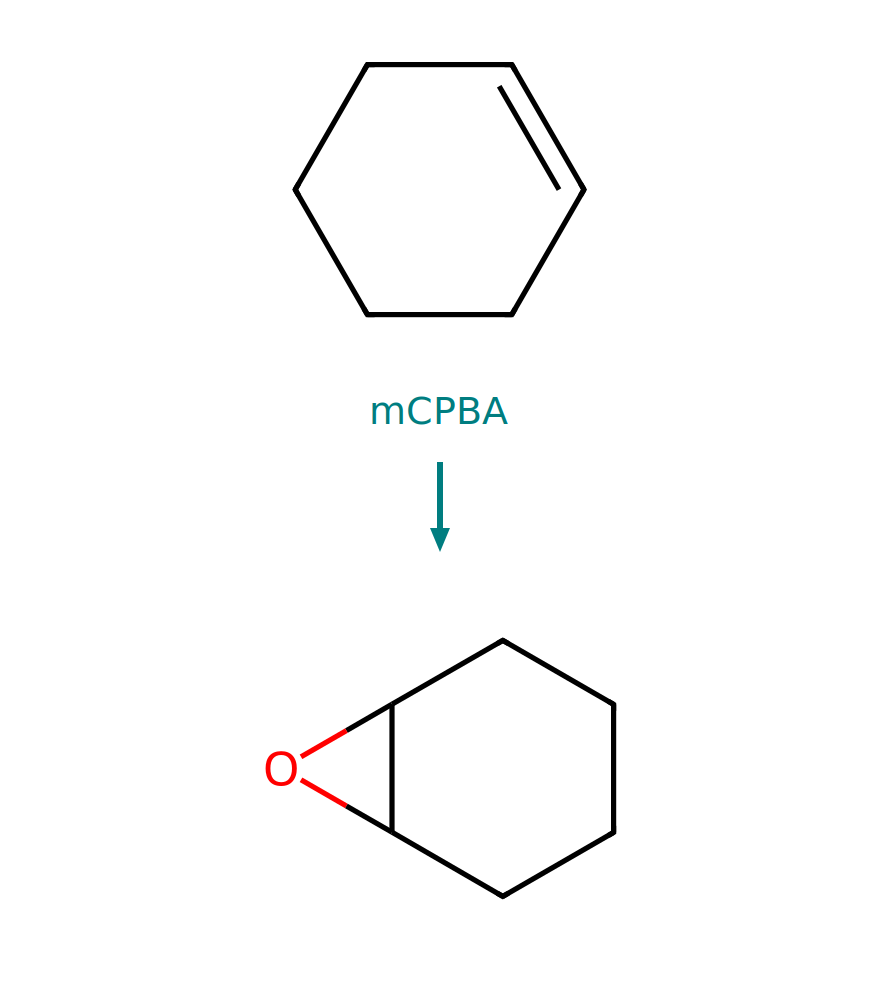
- A peroxyacid such as mCPBA transfers oxygen in a concerted epoxidation; the alkene’s relative geometry is retained in the epoxide.
- OsO₄ followed by suitable workup gives a syn vicinal diol. Epoxidation followed by aqueous acid opening gives an anti diol overall.
f. Ozonolysis
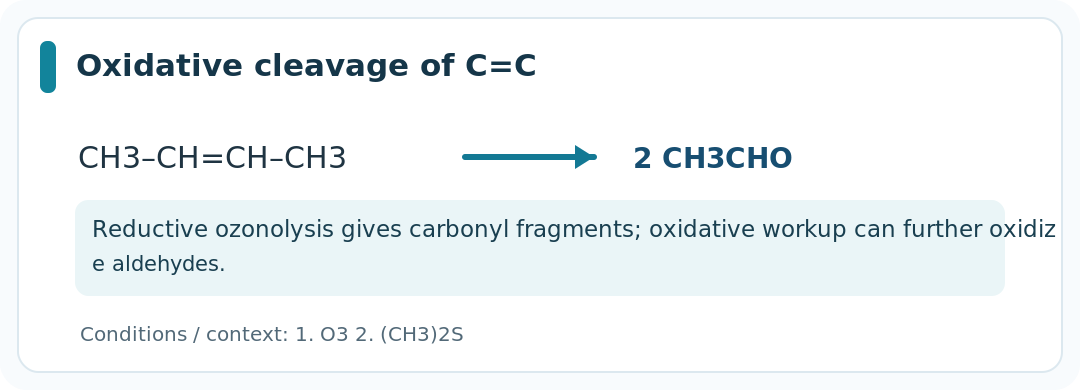
- O₃ followed by reductive workup, such as dimethyl sulfide, cleaves a C=C into carbonyl fragments. A carbon bearing H becomes an aldehyde; a carbon with two carbon substituents becomes a ketone.
- An oxidative workup can oxidize aldehyde fragments further to acids. Always state the workup before assigning final products.
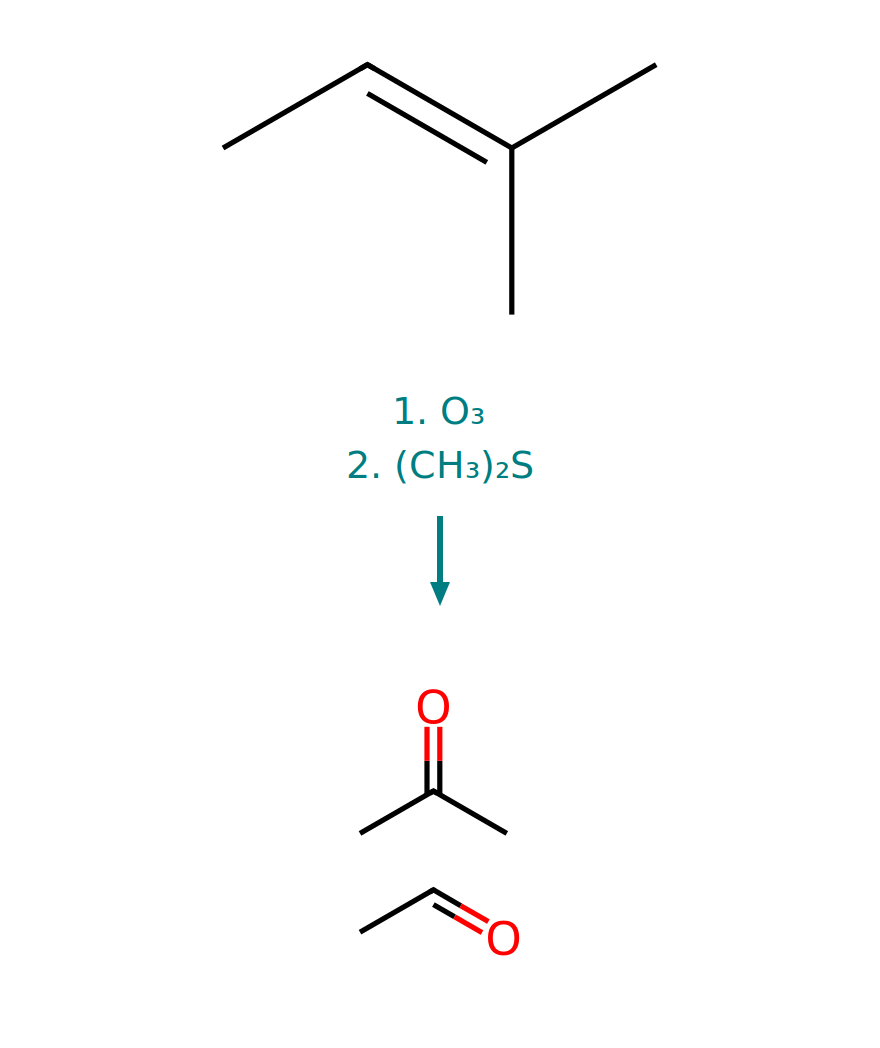
- Count carbon atoms in both carbonyl fragments to confirm conservation of the original alkene carbon skeleton.
g. Permanganate
- Cold dilute permanganate can give syn dihydroxylation; hot vigorous oxidative conditions can cleave the double bond.
- Under vigorous cleavage, an alkene carbon with H gives a carboxylic acid; a fully substituted alkene carbon gives a ketone; a terminal CH₂ carbon can be oxidized to CO₂.
h. Partial reduction
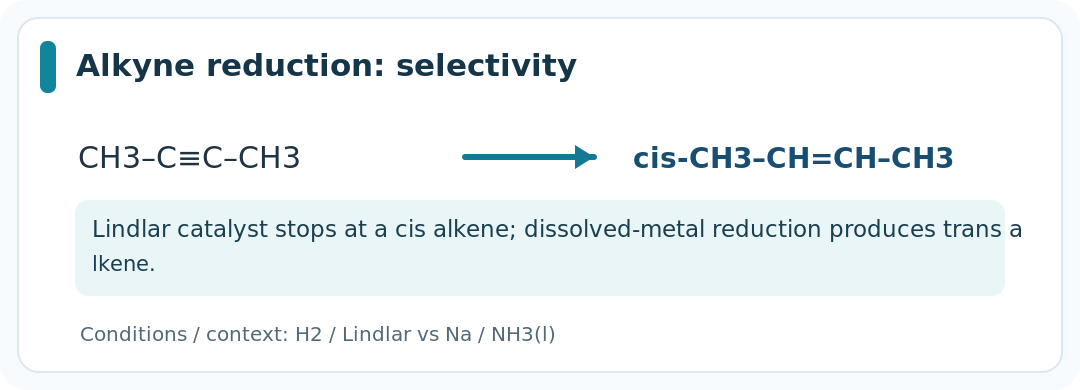
- H₂/Lindlar catalyst stops at the cis alkene for appropriate internal alkynes. Na in liquid NH₃ gives the trans alkene by dissolving-metal reduction.
- Excess H₂ with an active metal catalyst normally reduces the alkyne all the way to an alkane. Terminal alkene products do not have cis/trans geometry.
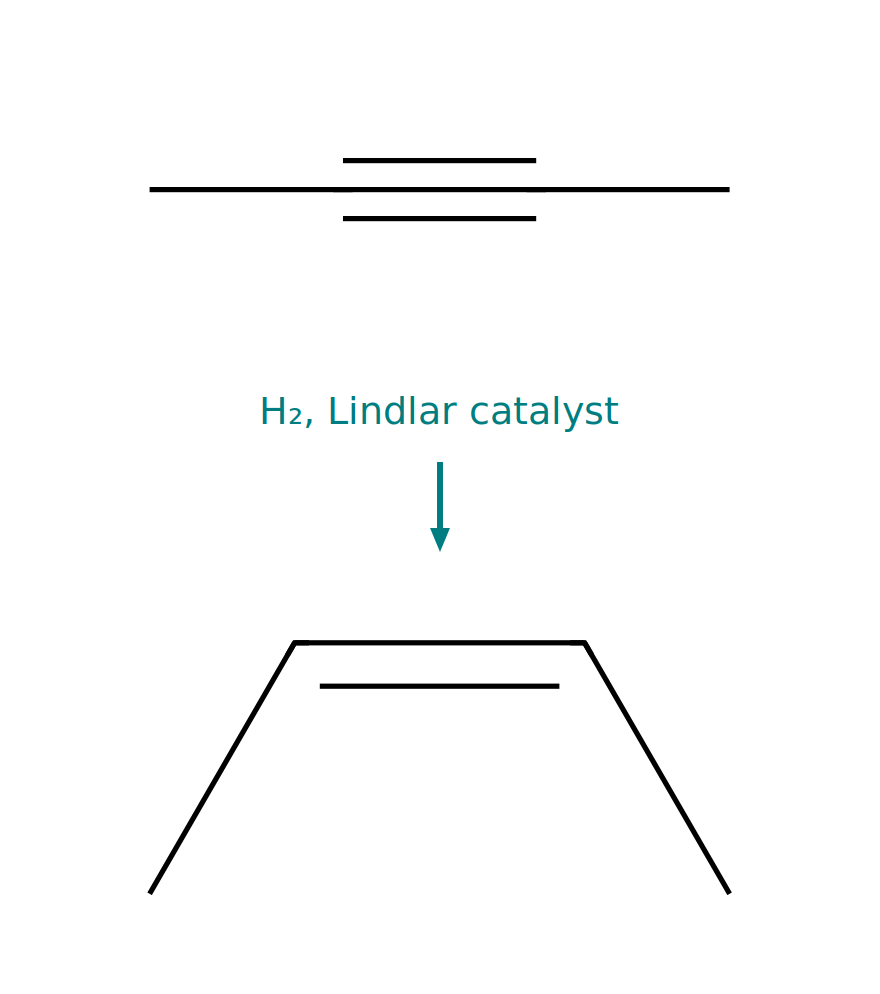
- Lindlar is a poisoned palladium catalyst; Na or Li in liquid ammonia can provide dissolving-metal partial reduction of suitable internal alkynes.
i. HX and X₂
- One equivalent of HX gives a vinyl halide; a second can give a geminal dihalide under ordinary Markovnikov conditions.
- One equivalent of X₂ gives a dihaloalkene, often with anti addition favored; excess X₂ can give a tetrahalide. Stoichiometry is essential.
| Reagent | Typical product |
|---|---|
| H₂, Lindlar | cis alkene |
| Na, NH₃(l) | trans alkene |
| Excess H₂, Pd/Pt/Ni | Alkane |
| 2 equivalents HX | Geminal dihalide |
- For a terminal alkyne under ordinary Markovnikov conditions, X generally bonds to the internal carbon. Conditions and rates need not match those of an alkene.

j. Alkyne hydration and carbonyl products
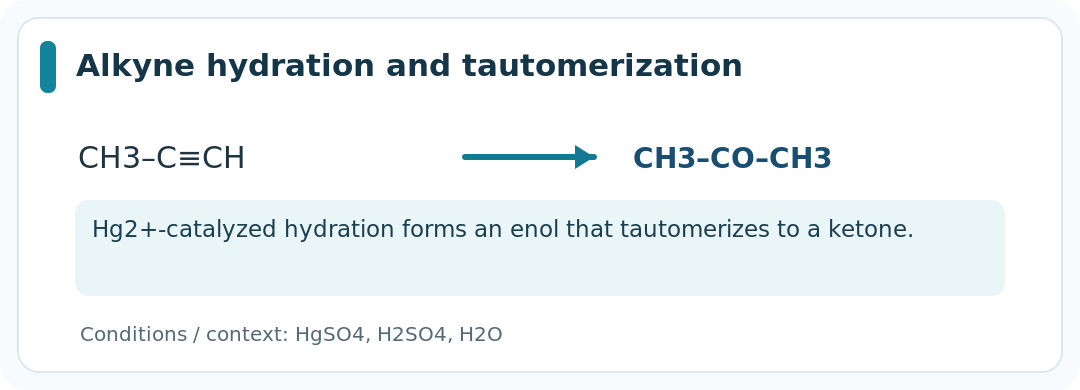
- HgSO₄/H₂SO₄/H₂O hydrates an alkyne through an enol that tautomerizes to a carbonyl compound. A terminal alkyne other than acetylene usually gives a methyl ketone.
- A bulky borane followed by H₂O₂/OH⁻ gives the other hydration orientation; a terminal alkyne gives an aldehyde after tautomerization.

- The initially formed enol undergoes proton transfer and double-bond relocation to the keto tautomer.
- For an internal unsymmetrical alkyne, the hydration regioselectivity needs substrate-specific analysis.

k. Acetylide chemistry
- NaNH₂ can deprotonate a terminal alkyne. The acetylide adds to aldehydes/ketones or substitutes at a suitable methyl/primary halide to form a C–C bond.
- Secondary/tertiary halides usually favor elimination with acetylide. Internal alkynes lack the terminal acidic H.
E. Conjugated Dienes
1. Conjugation, allylic reactions and cycloaddition
a. Adjacent π systems
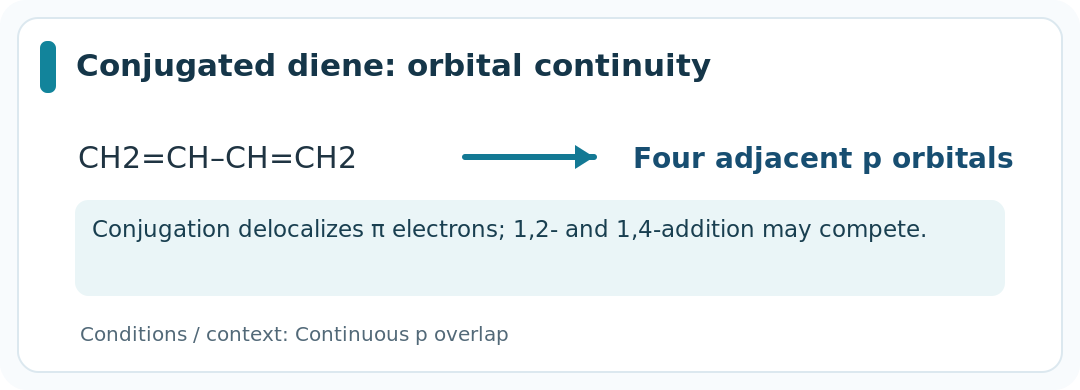
- A conjugated diene has alternating double and single bonds such as CH2=CH–CH=CH2.
- Four neighboring p orbitals permit electron delocalization over multiple atoms.
- Conjugated systems have different bond lengths and electronic behavior than isolated double bonds.
- Addition to a conjugated diene can give products from attack at the 1,2 and 1,4 positions; relative proportions depend on conditions.
b. 1,2- and 1,4-addition
- Conjugated dienes have alternating pi and sigma bonds and can form resonance-stabilized allylic intermediates. Electrophilic addition can give both 1,2- and 1,4-products.
- Kinetic and thermodynamic product preferences depend on barriers, product stability, temperature and reversibility; a product position alone does not define which control applies.
- NBS under radical conditions promotes allylic bromination while maintaining a low Br₂ concentration; resonance can lead to more than one allylic product.
c. Diels–Alder reaction
- A conjugated diene in an s-cis conformation reacts with a dienophile in a concerted [4+2] cycloaddition to make a six-membered ring.
- Relative cis/trans geometry on the dienophile is retained. Endo approach is often favored kinetically for suitable electron-withdrawing substituents, but is not an unconditional rule.
- Track the substituent positions and relative stereochemistry through the cycloaddition.
