
CH 3. Nomenclature, Conformations & Stereochemistry
A. Functional Groups and Naming
1. Representations, classes and nomenclature
a. Structural recognition
- A functional group is a characteristic arrangement of atoms that often controls chemical behavior.
- A carbon skeleton can be saturated or contain π bonds; functional groups containing oxygen, nitrogen, sulfur or halogens are identified by their connectivity.
- A single molecule can contain several functional groups, and each must be considered when predicting its properties.

b. Carbon framework and heteroatoms
- Alkanes contain only C–C and C–H single bonds; alkenes contain C=C and alkynes contain C≡C.
- Alcohols contain C–OH and ethers contain C–O–C. The oxygen of a carbonyl group is doubly bonded to carbon.
- Halides contain C–X (X = F, Cl, Br or I); amines contain a nitrogen bonded to carbon and/or hydrogen.
c. Substitution at carbon
- A carbon attached to one other carbon is primary (1°); one attached to two carbons is secondary (2°); one attached to three carbons is tertiary (3°).
- A carbon attached to four carbon atoms is quaternary (4°).
- The classification refers to the particular carbon atom, not the total size of the molecule.

d. Saturated hydrocarbons
- An acyclic alkane has the molecular formula CₙH₂ₙ₊₂. Carbon normally has four bonds, and a simple alkane carbon is approximately tetrahedral (sp³).
- Cycloalkanes with one saturated ring have formula CₙH₂ₙ. The ring removes two hydrogens relative to the corresponding acyclic alkane.
e. Constitutional isomers
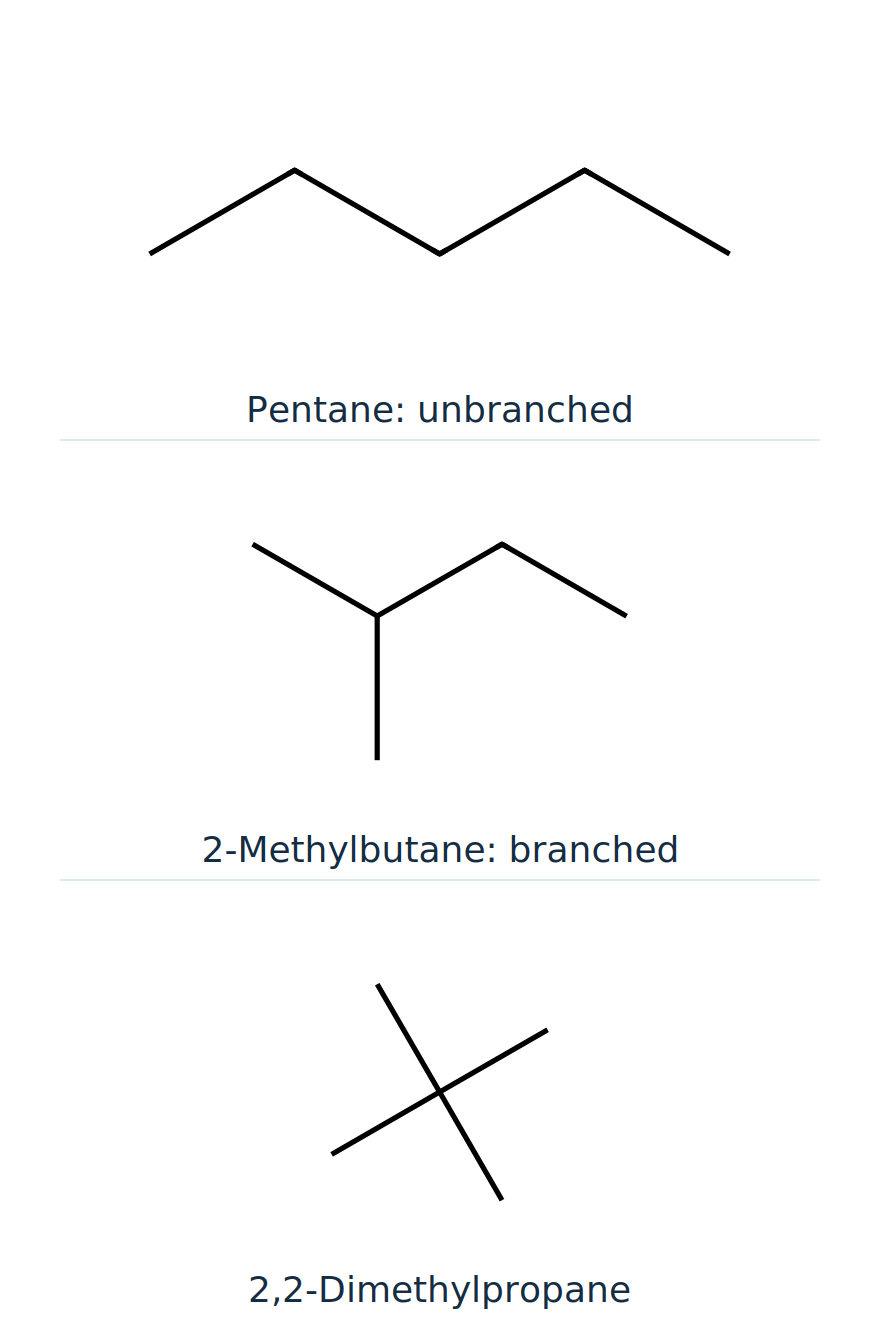
- Constitutional isomers have the same molecular formula but different atom connectivity.
- Pentane, 2-methylbutane, and 2,2-dimethylpropane all have the formula C₅H₁₂ but distinct carbon skeletons.
- A drawing flipped or rotated without changing bonds is not a different constitutional isomer.

f. Parent and locants
- Choose the longest continuous carbon chain for the parent name; where equal choices exist, apply substituent and locant rules.
- Number the chain to give the set of substituent positions the lowest possible locants, comparing at the first point of difference.
- Name branches as substituents and list different substituents alphabetically, ignoring multiplicative prefixes such as di- and tri-.

- Use parentheses and local numbering for complex substituents.
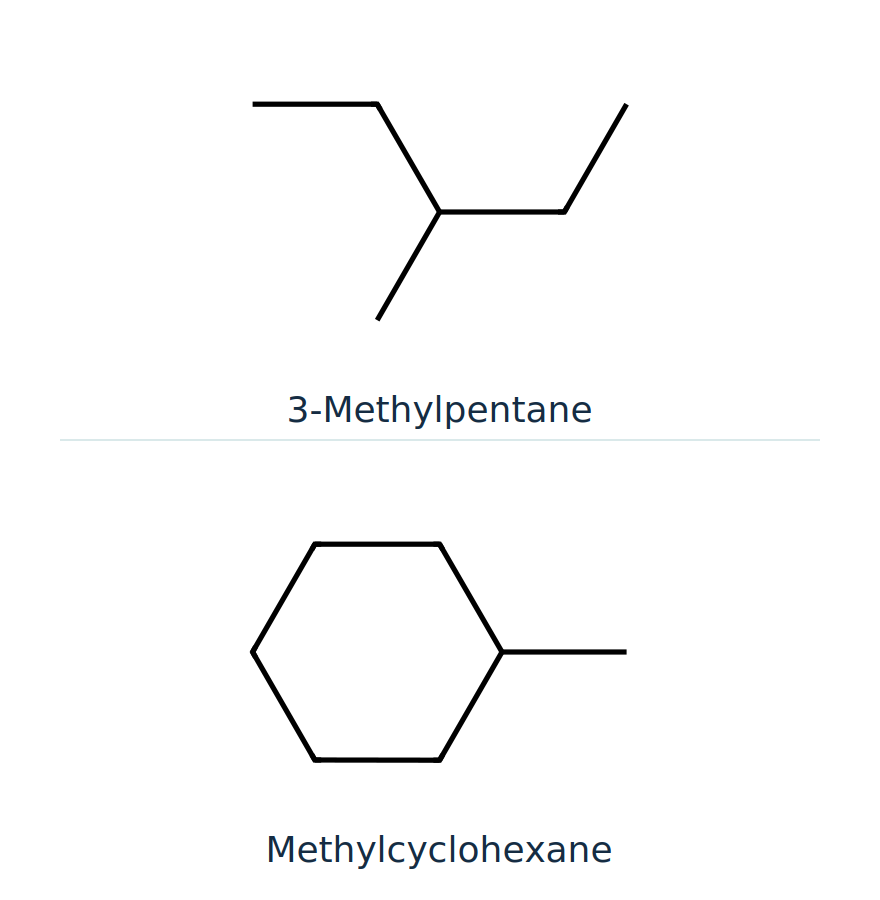
- Select a parent containing the principal functional group; its priority controls the suffix. Appropriate suffixes distinguish acids, esters, amides, aldehydes, ketones, alcohols, alkenes and alkynes.
g. Repeated substituents
- Use di-, tri- or tetra- for repeated identical groups and report each location.
- Separate numbers with commas and separate a number from a word with a hyphen.
- For example, CH₃–CH₂–CH(CH₃)–CH₂–CH₂–CH₃ is 3-methylhexane.
h. Condensed and skeletal notation
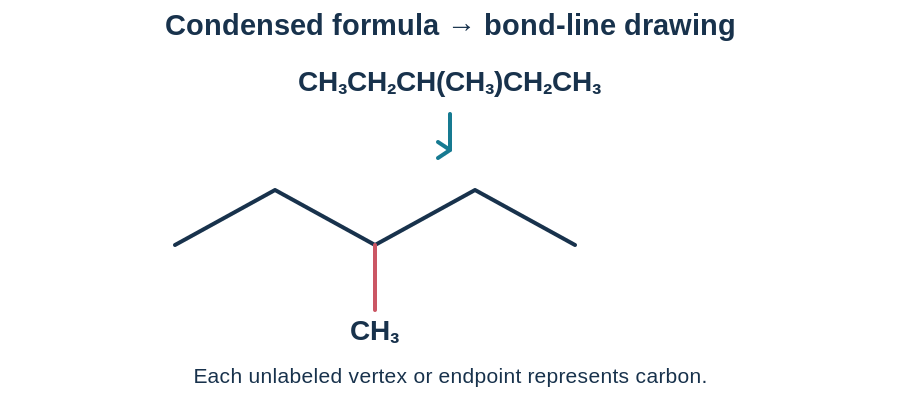
- Condensed formulas group bonded atoms in readable sequences; parentheses indicate a branch connected to the preceding atom.
- In bond-line structures, each unlabeled endpoint or vertex denotes a carbon and enough hydrogens are implied to satisfy normal valence.
- Heteroatoms and hydrogens attached to heteroatoms are normally shown explicitly.
i. Cyclic frameworks
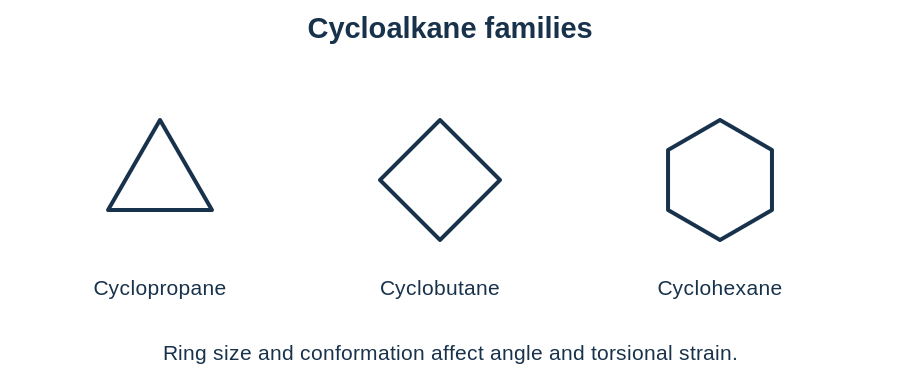
- Cycloalkanes are saturated hydrocarbons with at least one ring.
- Small rings experience a combination of bond-angle and eclipsing strain; cyclopropane is highly strained.
- Cyclohexane can adopt a low-strain chair conformation rather than a flat hexagon.
j. Rings and related structures
- For a simple substituted cycloalkane, compare the ring and attached chain when selecting the parent. Number ring substituents to give the appropriate lowest locants.
- Constitutional isomers have the same molecular formula but different connections. Conformers retain the same connections and differ by rotation or ring motion.
- Bicyclo[a.b.c] names count the atoms in each bridge between bridgeheads, excluding the bridgeheads; add two to obtain the total skeletal atom count.
| Relationship | What changes? |
|---|---|
| Constitutional isomers | Atom connectivity |
| Conformers | Torsion angles / ring shape |
| Configurational stereoisomers | Persistent spatial arrangement |
k. Naming
- Select the appropriate parent containing the multiple bond and assign its locant. Use -ene, -yne, -diene or -en-yne as appropriate.
- A terminal CH₂= group has two identical hydrogen substituents on one carbon and cannot display E/Z isomerism.

B. Newman Projections
1. Rotation, torsional energy and sterics
a. Single-bond rotation
- Rotation about a C–C σ bond creates conformations without changing bond connectivity.
- Conformations interconvert readily at room temperature unless restricted by a ring or severe steric constraints.
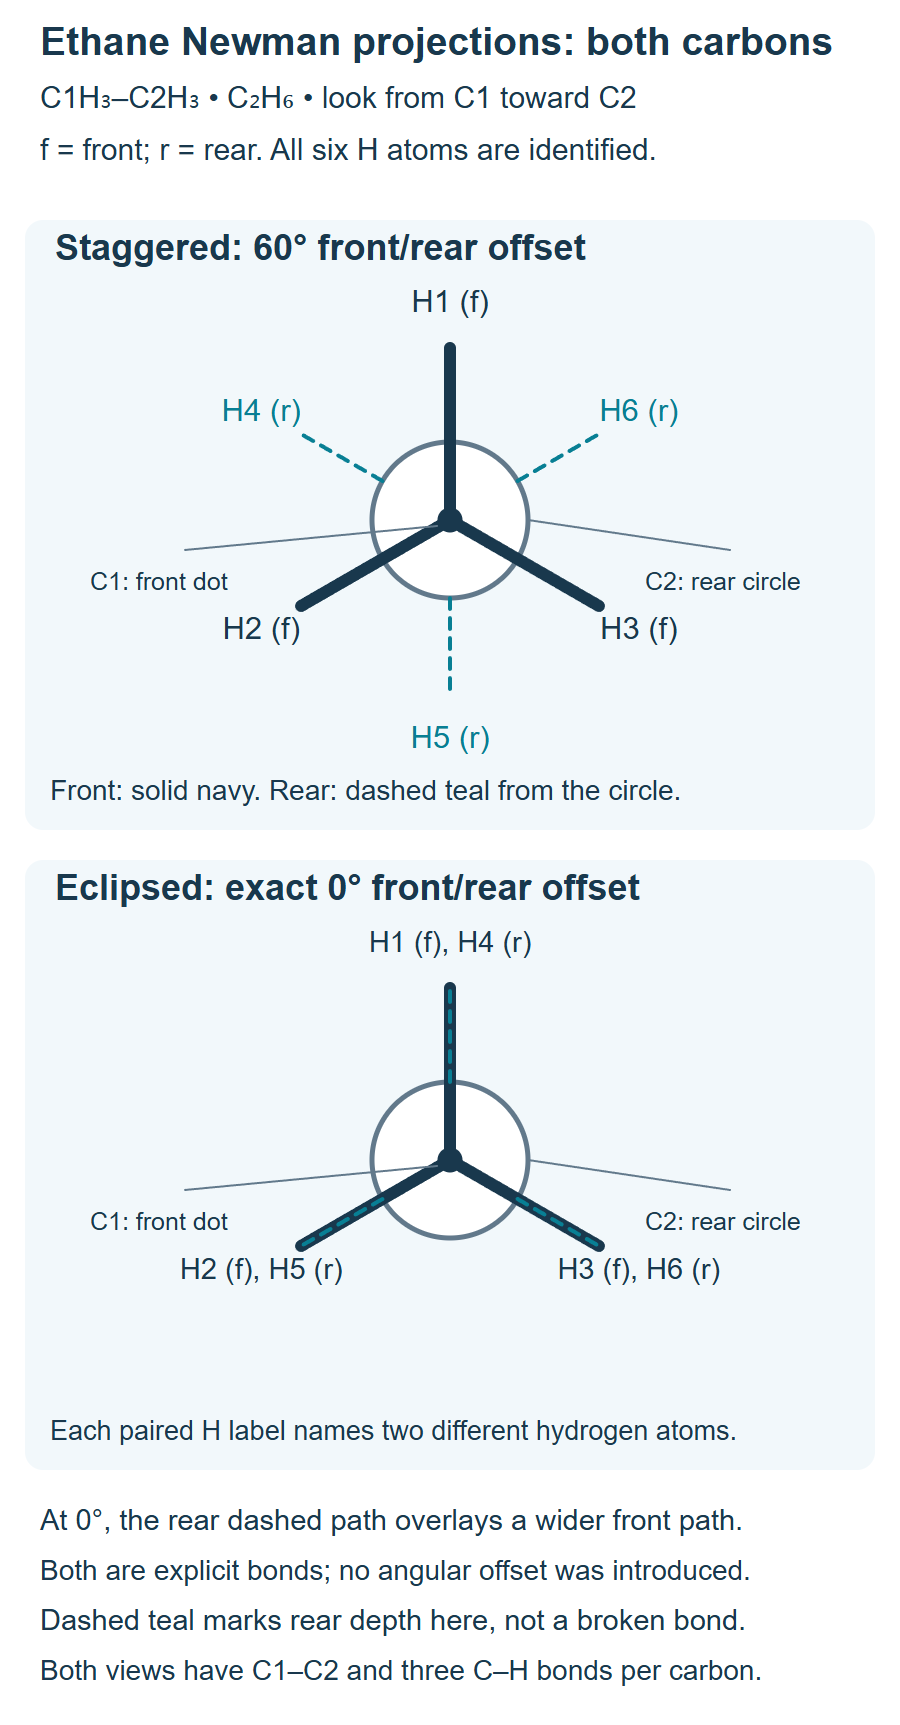
- Newman projections depict the positions of groups while looking directly along a C–C bond.
b. Staggered and eclipsed
- A Newman projection looks down a bond. The front atom is a point and the back atom a circle. Rotation changes the dihedral angle.
- Staggered ethane has lower energy than eclipsed ethane because eclipsing increases torsional strain.

- In staggered conformations, front and rear bonds are offset; in eclipsed conformations they align. For ethane, torsional energy changes periodically with the dihedral angle.

c. Butane conformations
- Anti butane places the methyl groups 180° apart and is the lowest-energy staggered conformer. Gauche places them 60° apart and has a steric penalty.
- An energy maximum is an eclipsed geometry; a minimum is a staggered conformer. Conformational interconversion normally does not require breaking a sigma bond.
C. Cyclohexane Chair Conformations
1. Ring geometry and conformational stability
a. Axial and equatorial bonds
- A chair cyclohexane has six axial and six equatorial C–H bond directions.
- Axial positions alternate up and down around the ring, and equatorial positions point roughly outward.
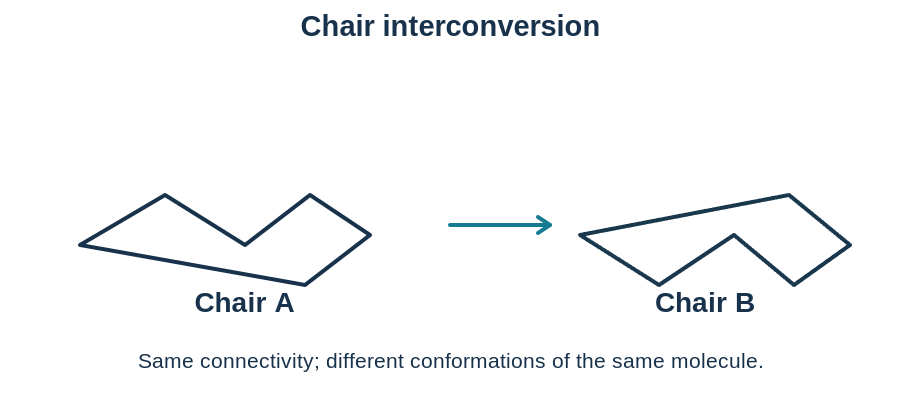
- A chair flip exchanges axial and equatorial positions while keeping each substituent on its original up or down face.
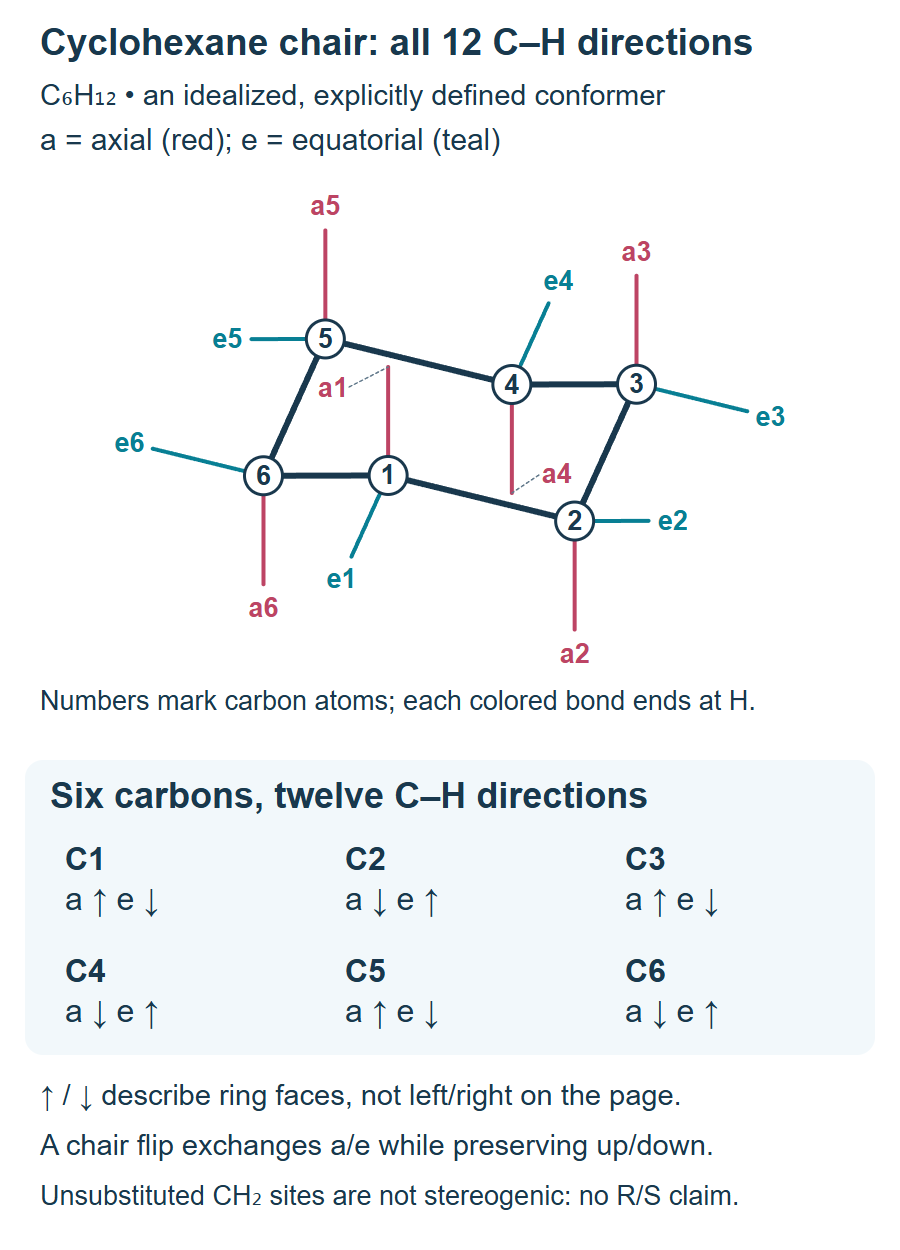
- A chair flip changes ring conformation rather than connectivity and preserves configuration at a stereogenic center. For a monosubstituted cyclohexane, the substituent exchanges axial and equatorial positions.
| Feature | Geometry | Interpretation |
|---|---|---|
| Axial | Approximately parallel to ring axis | Alternates up and down |
| Equatorial | Projects roughly outward | Often favorable for large substituents |
| Chair flip | Axial ↔ equatorial | Retains up/down configuration |
| Cis/trans | Relative ring face | Not altered by chair flip |
- For E2 in a cyclohexane chair, the reacting β-H and leaving group may need a trans-diaxial arrangement; the elimination mechanism is developed in CH 4.
b. Sources of strain
- Angle strain arises when bond angles depart from their preferred values; torsional strain comes from eclipsing. Steric crowding can add another contribution.
- Cyclopropane has severe angle strain; cyclobutane puckers, cyclopentane adopts nonplanar conformations, and cyclohexane strongly favors a chair.

c. Relative configuration
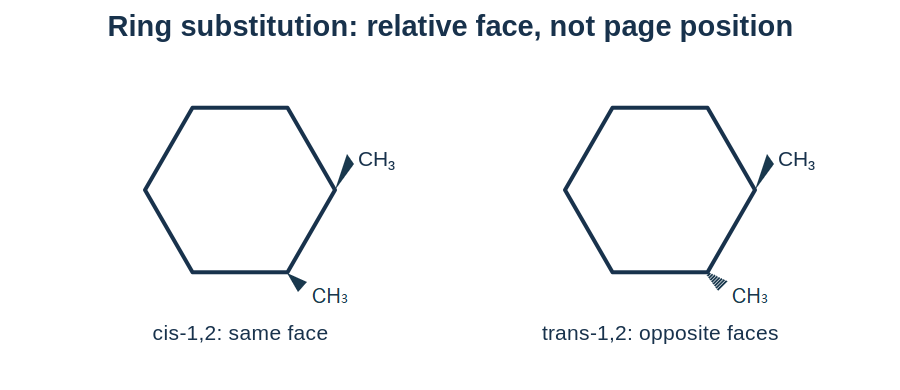
- Cis substituents lie on the same ring face; trans substituents lie on opposite faces. These relationships do not change during a ring flip.
- For two substituents, first mark up/down, then assign axial/equatorial in the chosen chair.

- Up/down ring-face orientation differs from axial/equatorial placement in a particular chair. A ring flip changes the latter but does not change cis/trans configuration.
d. Stability and fused rings
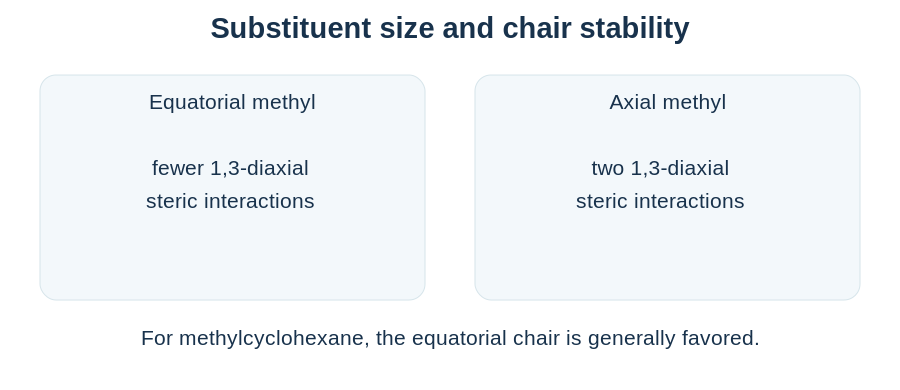
- Axial substituents experience 1,3-diaxial interactions. Compare total axial penalties, rather than counting only one substituent.
- For 1,2- and 1,4-disubstitution, trans can be diequatorial; for 1,3-disubstitution, cis can be diequatorial. Fused rings may restrict the accessible conformations.
| Positions | Diequatorial relationship |
|---|---|
| 1,2 | trans |
| 1,3 | cis |
| 1,4 | trans |
- Bulky substituents often favor equatorial placement because an axial substituent experiences destabilizing 1,3-diaxial interactions.
D. R/S, E/Z, Enantiomers and Diastereomers
1. Configuration, symmetry and optical properties
a. Connectivity and stereoisomer relationships
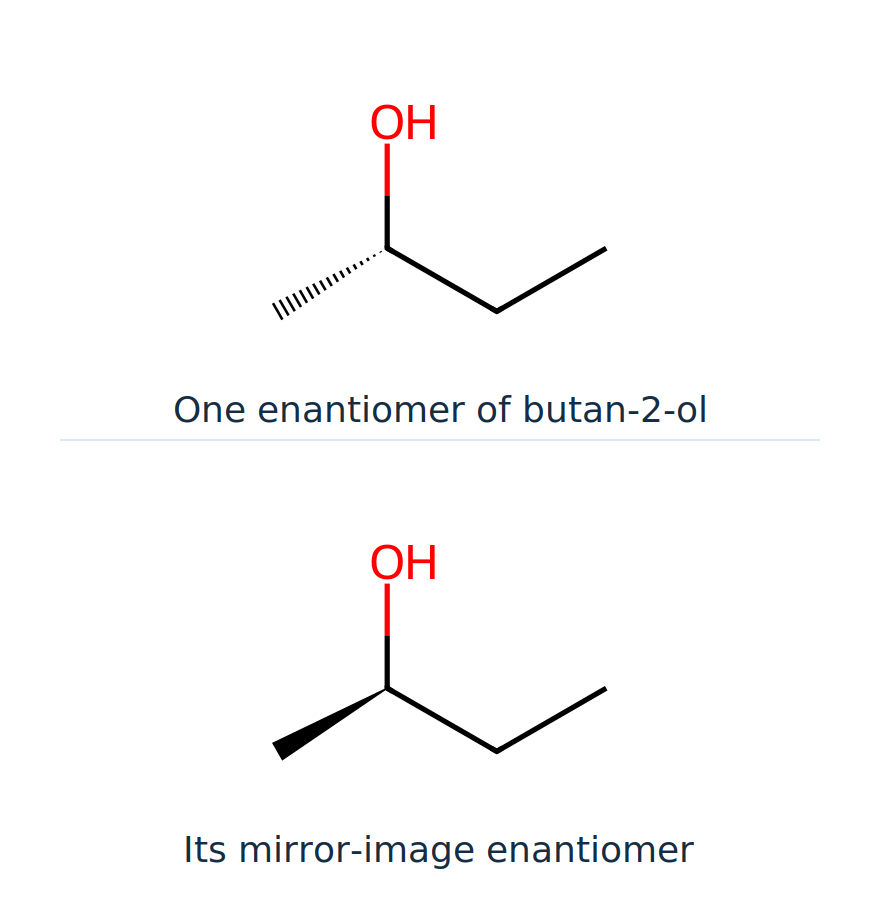
- Constitutional isomers have the same molecular formula but different atom connectivity. Stereoisomers retain connectivity and differ in spatial arrangement.
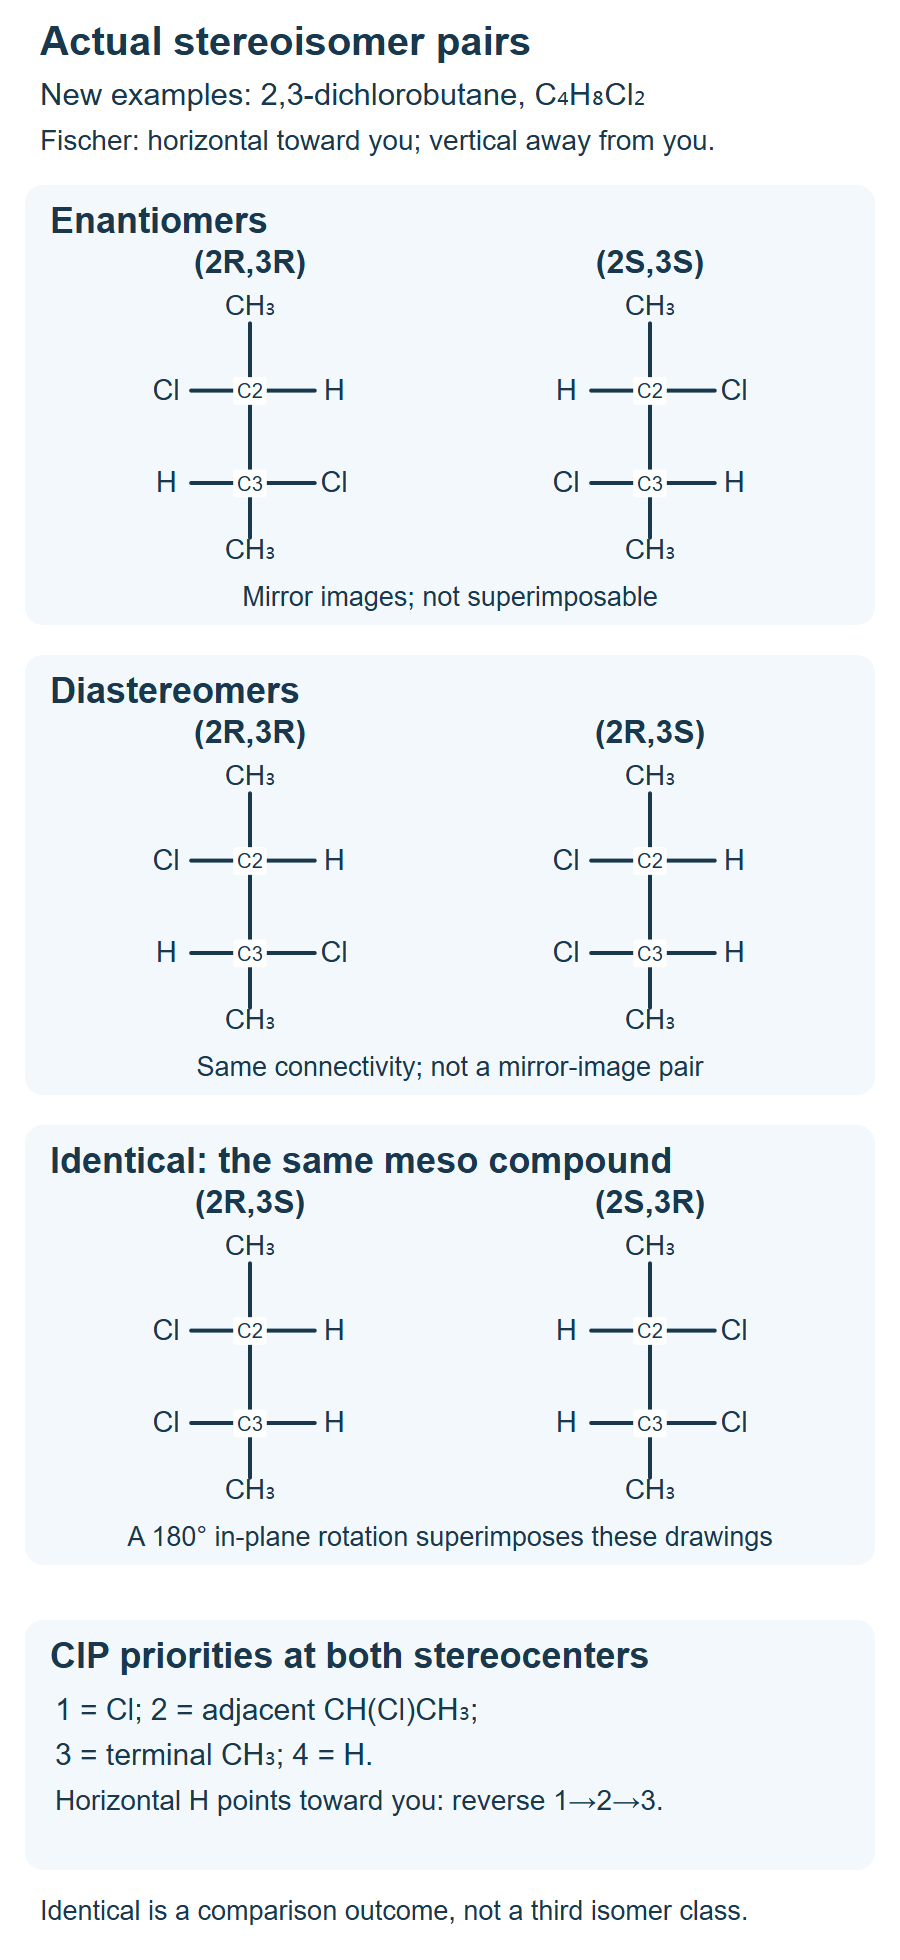
- Enantiomers are nonsuperimposable mirror-image stereoisomers. Diastereomers are stereoisomers that are not mirror images; suitable cis/trans and E/Z pairs belong to this category. Different spatial arrangements can give different physical properties.
b. Chirality, tetrahedral centers and molecular symmetry
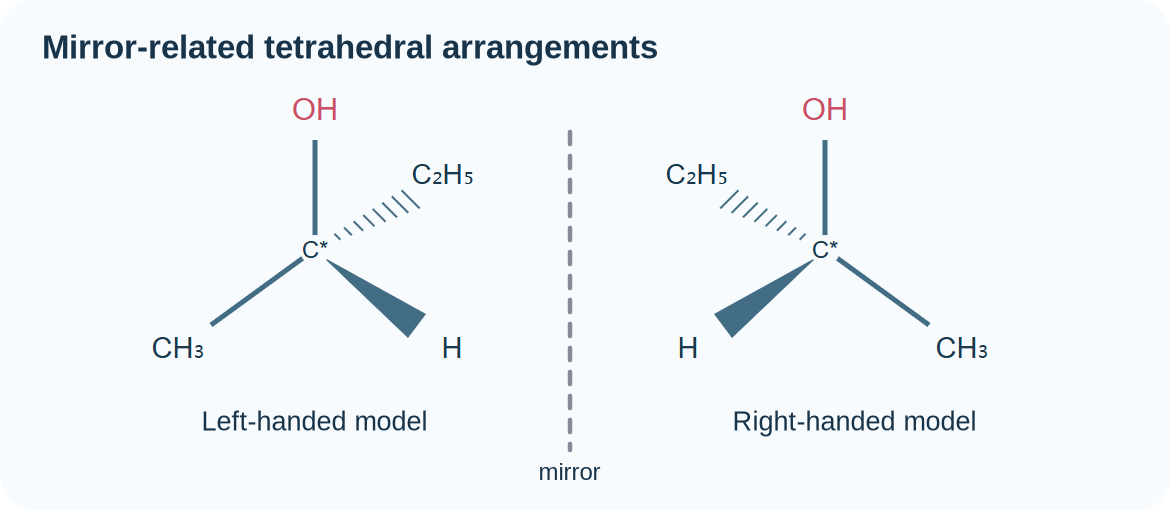
- A chiral molecule cannot be superimposed on its mirror image by rotation or translation; an achiral molecule can. Chirality is a property of the whole three-dimensional structure.
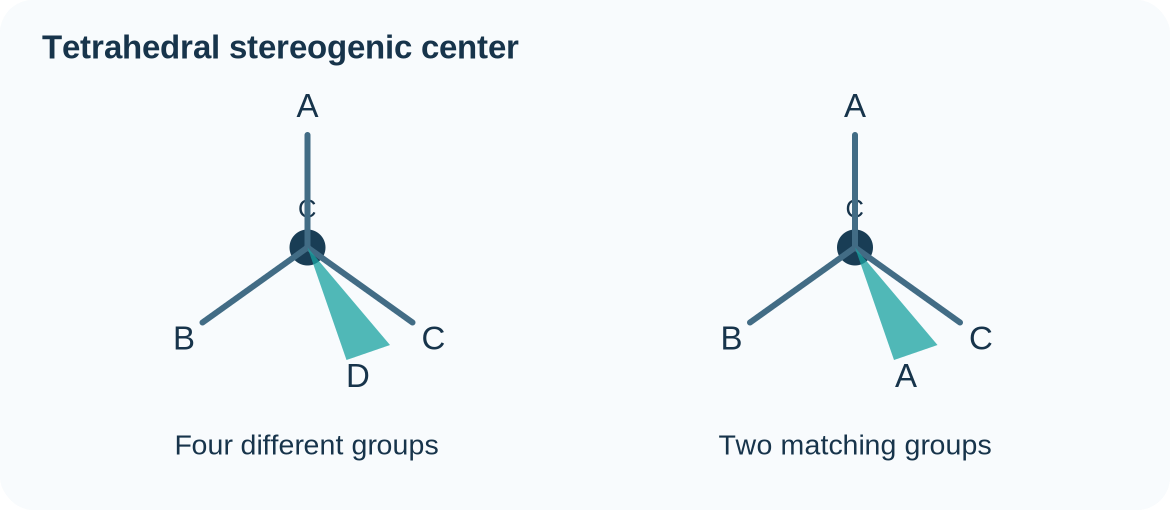
- A tetrahedral carbon with four different substituent groups is a common stereogenic center; one with two identical groups is not. A molecule with stereocenters can still be achiral through internal symmetry, and some chiral molecules have no tetrahedral carbon stereocenters.
| Relationship | Mirror images? | Superimposable? |
|---|---|---|
| Enantiomers | Yes | No |
| Diastereomers | No | No |
| Identical structures | May be | Yes |
c. Wedges and dashes
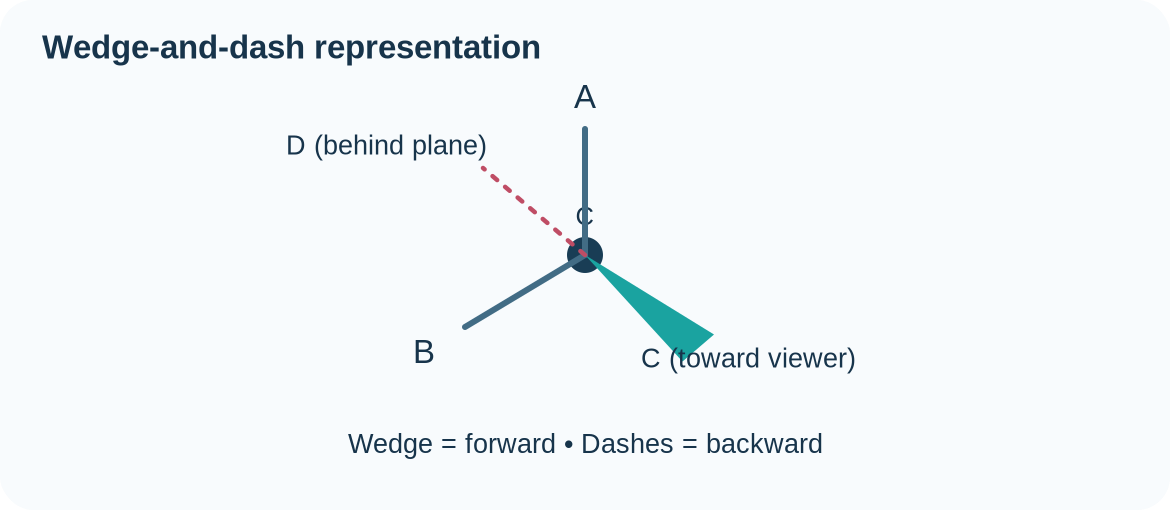
- A solid wedge represents a bond directed toward the viewer.
- A dashed bond represents a bond directed behind the page.
- Ordinary lines are drawn approximately in the plane of the page.
- Changing how a molecule is oriented on paper does not necessarily change its configuration.
d. First point of difference
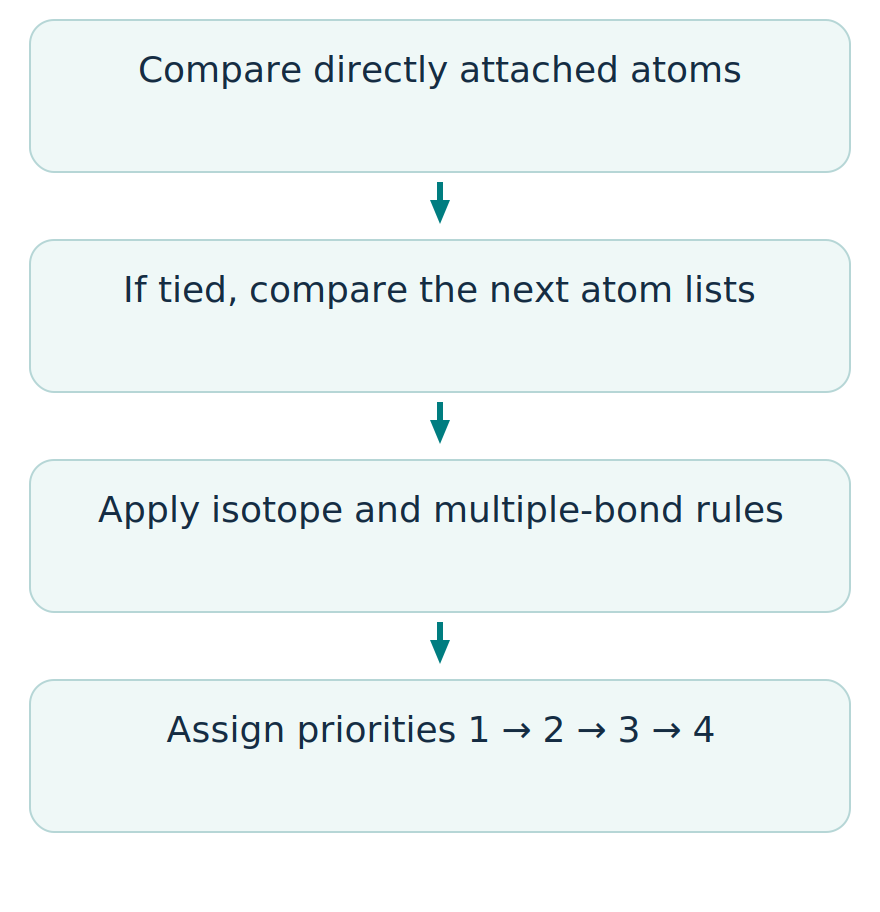
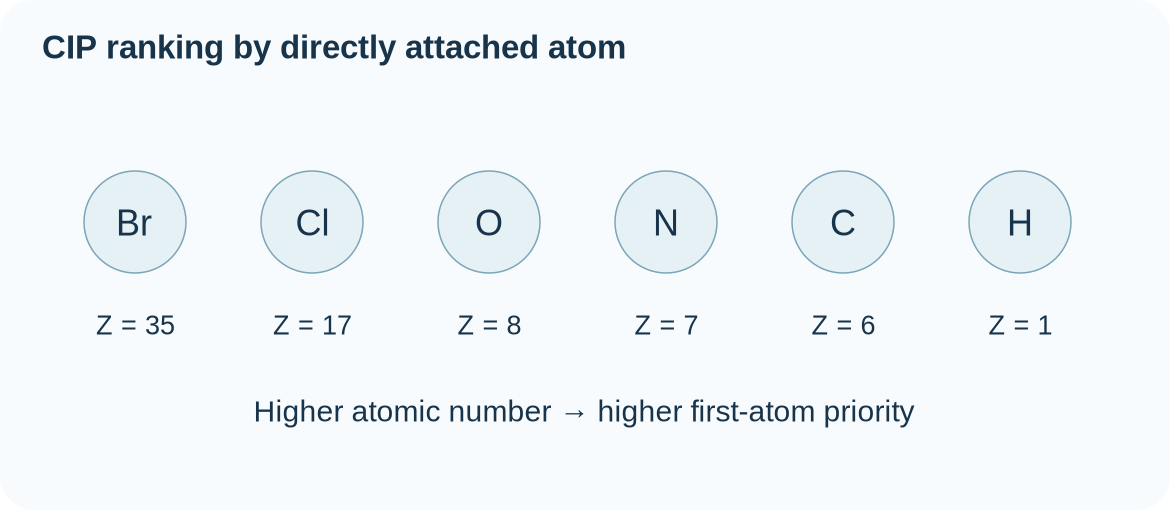
- Higher atomic number at the directly attached atom gives higher priority. For isotopes of the same element, higher mass number gives higher priority.
- At a tie, compare the next layer as ordered lists of attached atoms, starting from highest atomic number, until a difference appears.
e. Multiple bonds
- Treat multiple bonds using duplicate-atom conventions for the priority comparison. This is a ranking device, not a statement that actual duplicate atoms exist.
- A longer chain does not automatically outrank a shorter one; the first difference in the atomic-number comparison determines the result.
f. Orient the lowest priority
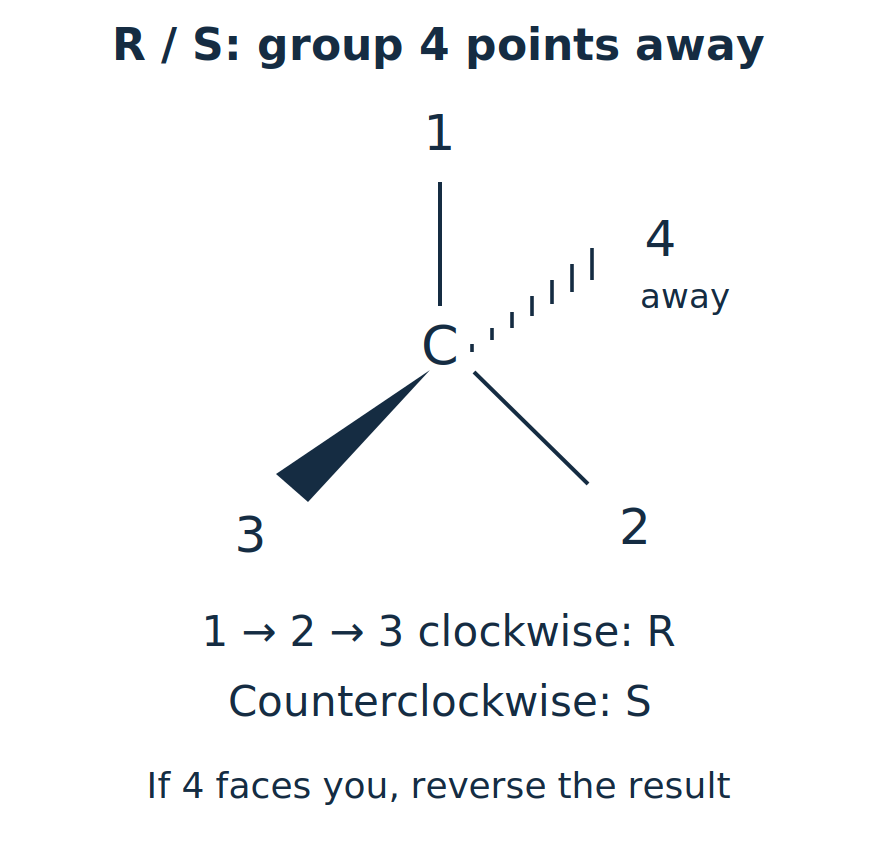
- Point group 4 away. Trace 1 → 2 → 3: clockwise is R and counterclockwise is S.
- If group 4 points toward the viewer, reverse the apparent result. A rotation of the whole molecule does not change configuration.

g. Permutation and labels
- Swapping any two groups reverses a single tetrahedral center; two such swaps restore its original configuration.
- R/S does not predict the sign of optical rotation. An R compound can be either (+) or (−).
h. Reading a Fischer projection
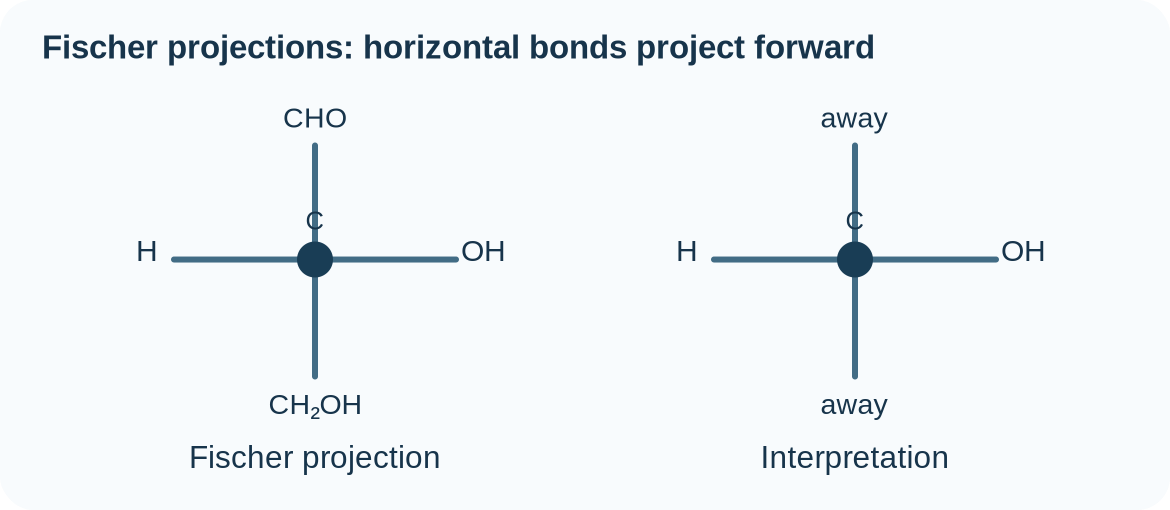
- The intersection of a Fischer projection represents a tetrahedral center. Horizontal bonds project toward the viewer; vertical bonds project away.
- Rotating the complete Fischer projection by 180° in the page preserves configuration; a 90° rotation generally changes it. Apply CIP priorities to the actual substituents before assigning R or S.

i. Assigning configuration
- Apply ordinary CIP priorities. If priority 4 is on a horizontal bond, reverse the apparent clockwise/counterclockwise assignment.
- Do not equate a vertical chain direction with a universal R/S label; compare the actual substituents at each center.
j. Geometric Isomerism
- Rotation around a double bond requires disrupting π overlap; geometrical arrangements can therefore remain distinct.
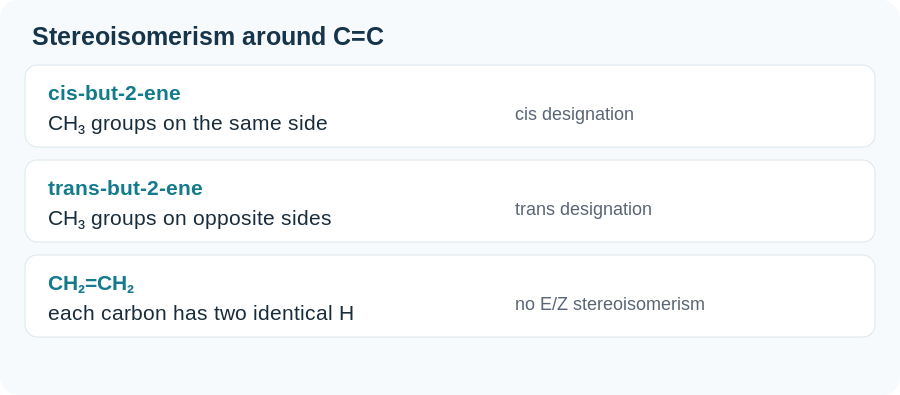
- For simple appropriately substituted alkenes, cis refers to corresponding groups on the same side and trans to opposite sides.
k. Requirement for E/Z
- Each carbon of the double bond must have two different attached groups for E/Z stereoisomerism to exist.
- A terminal CH₂= group cannot display E/Z because one carbon has two identical hydrogen atoms.
l. E and Z
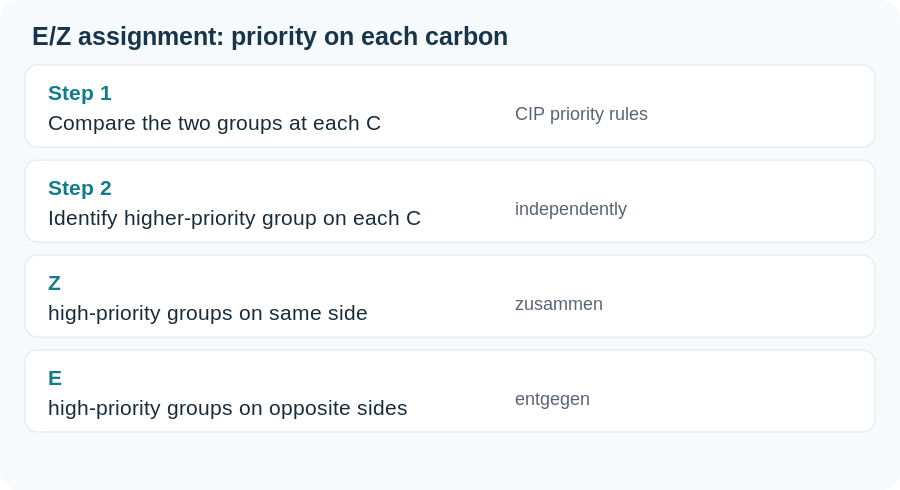
- Rank the two groups attached to each alkene carbon using CIP rules. Higher atomic number wins; at a tie, compare successive attached-atom lists.
- High-priority groups on the same side give Z; opposite sides give E. Cis/trans and E/Z describe different comparisons and should not be interchanged automatically.
- Assign priorities from substituent identity, not from arbitrary top/bottom positions in the drawing.
m. Stereoisomer counting and symmetry
- For n independent tetrahedral stereocenters, 2ⁿ is an upper limit on configurational stereoisomers. Symmetry, meso forms and equivalent configurations can reduce the count; count distinct structures rather than drawing duplicates.
- Inverting every center can produce an enantiomer, while inverting only some often gives a diastereomer. Molecular identity and symmetry must still be checked.

n. Meso symmetry and optical inactivity
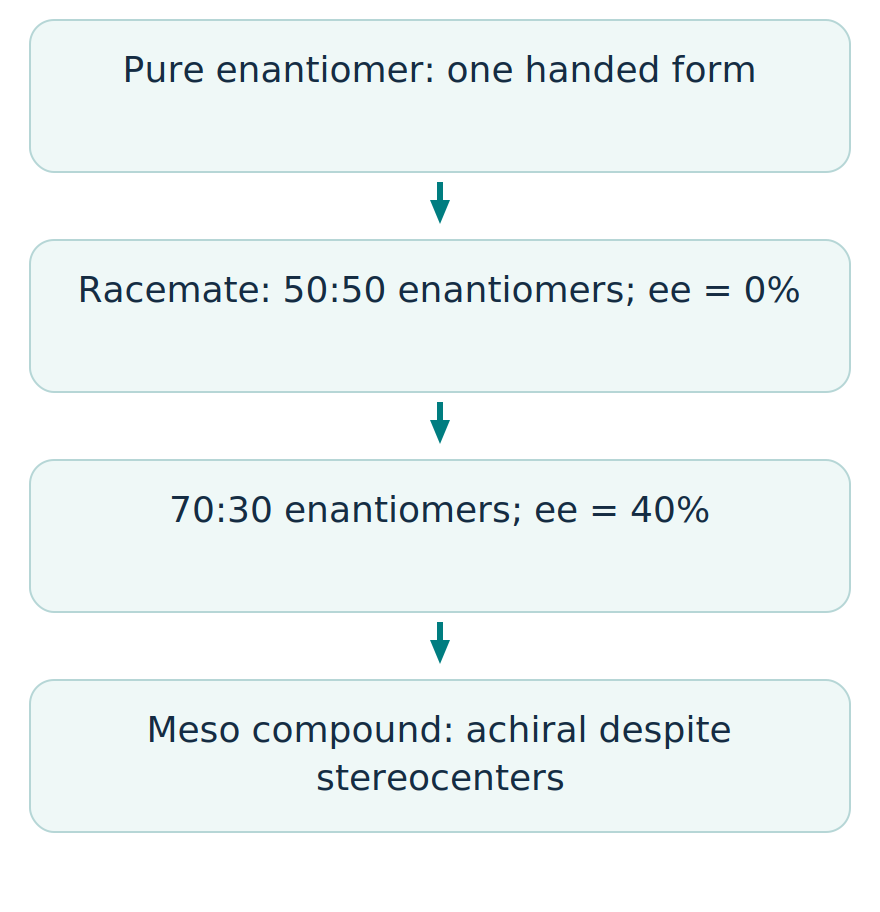
- A meso compound contains stereogenic centers but is achiral because its overall structure has the required internal symmetry. The stereocenter count alone cannot establish molecular chirality; internal symmetry can reduce the number of distinct stereoisomers below 2ⁿ.
- A pure meso substance is optically inactive through molecular achirality. A racemate is optically inactive because the equal and opposite rotations of its enantiomers cancel. These are different explanations.

o. Specific rotation
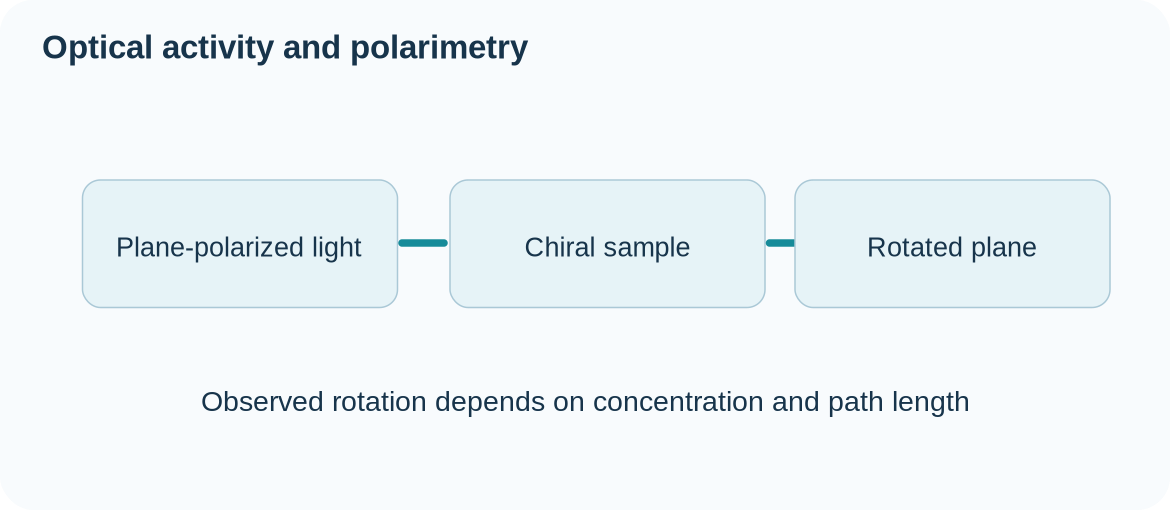
- Specific rotation normalizes observed rotation to path length and concentration under specified wavelength, temperature, solvent and concentration conventions.
- Enantiomers rotate plane-polarized light equally and oppositely under identical conditions. Diastereomers have no such simple equal-and-opposite relationship.
- Under the common solution convention, [α] = αobserved/(l × c), with l in dm and c in g/mL; confirm the concentration convention used by a reference.
- The sign of rotation, (+) or (−), does not determine an R/S descriptor. Measured rotation depends on concentration, path length, temperature and wavelength.
p. Composition
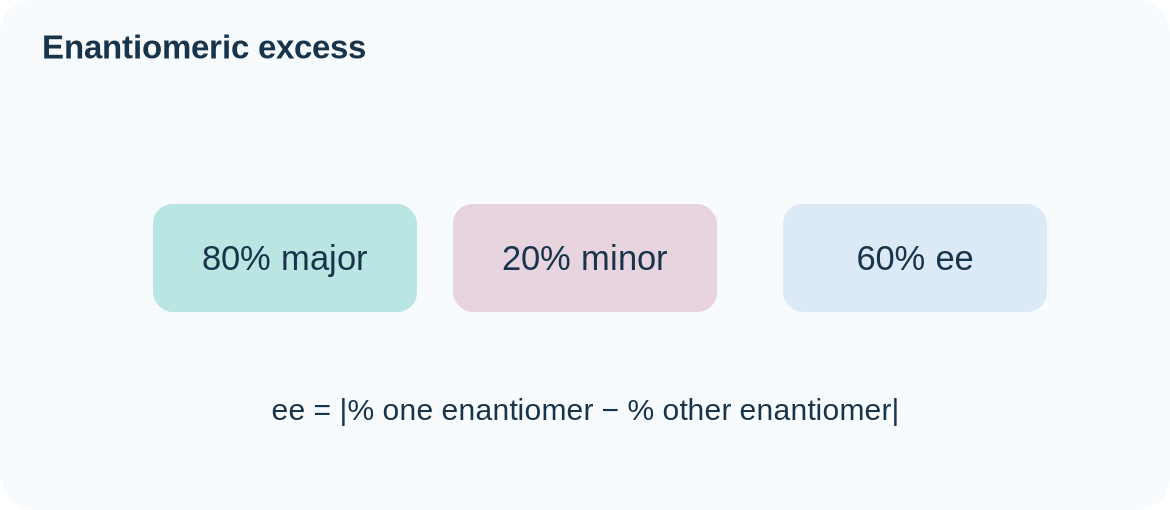
- For a mixture containing only two enantiomers, ee = |% major − % minor|. A 70:30 mixture has 40% ee.
- Using rotation to estimate ee requires the pure-enantiomer reference and compatible measurement conditions. A zero rotation alone does not prove that a sample is achiral or pure.
- A racemate contains equal enantiomer amounts and has zero net rotation under usual conditions. An 80:20 enantiomer mixture has 60% ee.
q. Chiral recognition
- Enantiomers have identical properties in many achiral environments, apart from opposite optical rotation and interactions with polarized light.
- A chiral reagent, catalyst, receptor, or stationary phase may distinguish between two enantiomers.
- Biological interactions are frequently stereoselective because biomolecules are chiral.
r. Configuration versus optical rotation
- A reaction can retain or invert a configuration at a stereogenic center, depending on the mechanism.
- R/S descriptors may change when the substituent priorities change, even if the three-dimensional arrangement is retained.
- A racemic or enriched product distribution depends on reaction pathway and stereochemical environment.